The Evolutionary Persistence of Male Androphilia: A Causal-Portfolio Theory of Developmental Paths and Selection Accounting
Department of Psychiatry, Perelman School of Medicine, University of Pennsylvania; Lifespan Brain Institute, Penn Medicine and Children's Hospital of Philadelphia
Download PDF manuscript and supplementary material, one file
Abstract
The conventional evolutionary paradox of male homosexuality is mis-specified because it combines distinct developmental and population-genetic accounting problems. Etiological heterogeneity does not remove the selective burden carried by inherited liability; it determines which burden belongs to which causal component. This article advances a causal-portfolio theory in which enduring male androphilia is defined by repeated male-directed and female-directed attraction rather than by identity, partnership, or behavior. Four definitions fix the explanandum, five causal levels, and one reporting scale. Four empirical propositions are placed at risk: androphilia is a convergent phenotype produced by at least two mechanism-distinct prenatal paths; its inherited architecture is distributed and shared across sexes; prenatal organization, not a scalar androgen deficit, is its principal causal stage; and its maintenance is component-specific, with inherited liability compensated in aggregate across full-genome partitions covering at least 60% of common-SNP heritability, the verdict confined to that covered majority, and recurrent prenatal exposures costly to remove. A path qualifies only if its standardized effect on latent attraction is at least .10 with a lower 90% bound above .05 in an independent hold-out, it survives an assumed residual mediator-outcome correlation of .20, and a design independent of sequential ignorability supports it. The core is rejected only when fewer than two paths qualify and a route-free comparator is equivalent to the best path model in a pooled hold-out; the operating characteristics of that decision are reported. Six critical studies separate failure of the core from failure of a subarchitecture, path, or assay.
Keywords: Male androphilia; Sexual orientation; Causal inference; Evolutionary maintenance; Falsifiability
Introduction
Male homosexuality is evolutionarily consequential because enduring androphilia has often reduced direct reproduction, shows familial and genetic influence, and nevertheless recurs across populations and historical periods (Apostolou, 2022; Bailey & Pillard, 1991; Bailey et al., 2000; Kendler et al., 2000). The burden is real. What is wrong is the assumption that the phenotype, its causes, and its persistence can all be represented by one inherited character at one equilibrium. That formulation demands one answer where the biology presents several different accounting problems. Throughout this article, androphilia names the repeated attraction construct in Definition 1. When a cited study measured homosexuality, same-sex behavior, identity, partnership, or legal union, its own phenotype term is retained and is not treated as interchangeable with androphilia.
Four questions must be separated. First, which inherited variants or polygenic components raise liability, and why do they remain polymorphic? Second, which maternal or fetal developmental systems can generate the phenotype, and why are the relevant vulnerabilities not eliminated? Third, why do noninherited pregnancy-dependent events recur in successive generations? Fourth, how does the joint distribution of inherited liability, maternal physiology, fetal exposure, and developmental regulation generate the observed prevalence in a particular cohort? Heterogeneity divides these burdens; it does not erase any of them.
The correct opening claim is therefore exact: the conventional paradox is mis-specified because it combines several distinct evolutionary and developmental accounting problems. An inherited component that substantially raises the probability of reduced reproduction still requires a population-genetic explanation. A recurrent maternal immune exposure does not. It instead requires an incidence model and an explanation for the persistence of the causal immune specificity, placental transfer process, fetal target, or downstream sensitivity. A theory that merely lists both processes has not solved either one.
The unit of inherited accounting is the variant or molecular component and its average fitness effect across every body in which it occurs. Female phenotypes remain inside that ledger because men and women share much of the same inherited architecture. The largest behavioral GWAS estimated a substantial cross-sex genetic correlation, approximately .63, even though expression differs by sex (Felesina, 2026; Ganna et al., 2019). The developmental explanandum can be male-specific without the evolutionary bookkeeping becoming male-only.
The theory also makes no group-selection claim. Coalition effects, pair bonding, kin investment, and social organization matter only when they change the survival or reproduction of carriers or genetic relatives and can be attributed back to inherited variation. Comparative evidence can show that same-sex behavior is common, plastic, or socially functional; it cannot by itself explain the persistence of stable human male-directed attraction.
The Causal-Portfolio Theory
Stated plainly, the theory holds that enduring male androphilia is a single endpoint reached by more than one developmental path. Inherited liability, maternal physiology, pregnancy history, and developmental chance act through prenatal endocrine, placental, immune, and regulatory mechanisms; those mechanisms organize how male and female cues are valued; and the resulting attraction profile is what behavior, identity, and partnership imperfectly express. Because the paths differ in kind, their persistence must be explained in different currencies. An inherited component is answerable to allele-level selection summed over every carrier of either sex; a recurrent pregnancy-dependent event is answerable to an incidence model and to the cost of removing the vulnerability it exploits. The theory is a portfolio in exactly that sense: the phenotype is produced by several holdings, each kept on its own ledger, and the maintenance verdict is rendered on the measurable aggregate of the inherited holdings rather than on any one favorable component. The four questions above are divided accordingly: the first falls to the inherited arm of P4; the second and third to its recurrent arm, which pairs a constraint test with an incidence model; and the fourth to the joint-distribution definition D4, while P1 through P3 state what the developmental architecture must be for that division of burdens to be the right one.
The theory is stated as four definitions and four empirical propositions. The distinction is deliberate and is not a matter of presentation. The definitions fix what is being explained, what counts as a route, and on which scale quantities are reported; they can be judged useful or useless, but they cannot be false. The empirical propositions can fail, and the critical studies below state how. Table 1 collects every prespecified quantity, threshold, and margin in one place; the text refers to Table 1 rather than restating the definitions at each use.
| Quantity | Prespecified value | Scale or basis |
|---|---|---|
| Explanandum region R | ĀM ≥ +1.80 and ĀF ≤ −0.75 | Male-population standardized longitudinal means |
| Measurement rule m | Person-level means over T ≥ 4 occasions spanning ≥ 5 years, then region R | Declared deterministic function of the latent summaries |
| Planning prevalence π | .025 (sensitivity .017 to .036) | Implied by R under the planning joint distribution |
| Planning attraction correlation | −.55 (sensitivity −.45 to −.65) | Bivariate-normal planning approximation for (ĀM, ĀF) |
| Materiality threshold | Standardized effect ≥ .10 | Latent attraction |
| Lower-bound gate | Lower 90% bound > .05 in an independent probability-weighted hold-out | Latent attraction |
| Robustness value ρ* | ≥ .20 | Assumed absolute residual mediator-outcome correlation |
| Independent identification | At least one design not resting on sequential ignorability | Within-family or within-mother contrast, instrument, targeted perturbation, or negative control |
| Practically null | < .03 | Latent attraction |
| Indeterminate zone | .03 to .10 | Latent attraction |
| Primary causal equivalence margin | .03 | Latent attraction; two one-sided tests at 90% confidence |
| Derived prevalence margin | .0015 under the planning assumptions; recomputed from the fitted joint distribution | Image of the .03 causal margin under m; dπ/dδ = .048 |
| Secondary predictive margins | ΔR² < .0025; ΔBrier < .002; Δ log-loss < .005 | Held-out prediction; secondary, never decisive alone |
| Rejection rule | Fewer than two qualifying paths and equivalence on the causal margin plus at least three of four secondary margins | Pooled probability-weighted hold-out |
| Stratification | Four reference-projected genetic-similarity strata; self-reported ancestry retained as a covariate and for invariance testing | Heterogeneity diagnostics, not gates (National Academies of Sciences, Engineering, and Medicine, 2023) |
| Consequential bound, Critical Study 3 | .10 with lower 90% bound above .05 | Identical to the materiality gate |
| Coverage criterion C | ≥ .60 with lower 95% bound ≥ .50 | Fraction of out-of-sample common-SNP heritability in selection-valid partitions |
| Planning h²SNP | .15 (planning range .08 to .25); estimated in the discovery sample | Repeated attraction; the coverage denominator is estimated, not assumed |
| Compensation criterion | Lower bound on GC/KC ≥ 1.0, or a separately fitted nonnegative equilibrium | Covered partitions only; no imputation to 1 − C |
| Planning design, Critical Study 1 | 10,000 analyzed men per stratum (40,000 total); 30% hold-out | Smallest grid point meeting the operating characteristics below |
| Target operating characteristics | ≥ .90 correct retention; ≥ .80 correct rejection; familywise false-decision probability ≤ .05 | End-to-end Monte Carlo simulation of the complete decision |
Note. All thresholds are fixed before data collection; where the text repeats a value, this table is authoritative. Standardized effects refer to mediated or identified edge-intervention effects on latent attraction. The robustness value is the smallest assumed absolute residual mediator-outcome correlation at which a path's lower 90% bound falls to .05.
Definitions and Commitments
D1. The explanandum is enduring male-directed attraction, not a broad social category. Attraction to men and attraction to women are measured separately and repeatedly. Fantasy, behavior, identity, partnership, and legal union are downstream or imperfect indicators. Predominant or exclusive male androphilia is the prespecified region R of the joint attraction distribution given in Table 1 and shown in Figure 2, not a synonym for any single observed indicator.
D2. Phenotypic distinctness and genetic separability are different claims. Orientation, behavior, identity, and partnership can remain phenotypically noninterchangeable even if one genetic factor contributes to all of them. The separate genetic claim, that cue-specific attraction and general behavioral propensity retain distinguishable inherited components after reliability and cross-sex covariance are modeled, appears as P2 and is tested in Critical Study 2. Failure of P2 does not make the observed phenotypes identical.
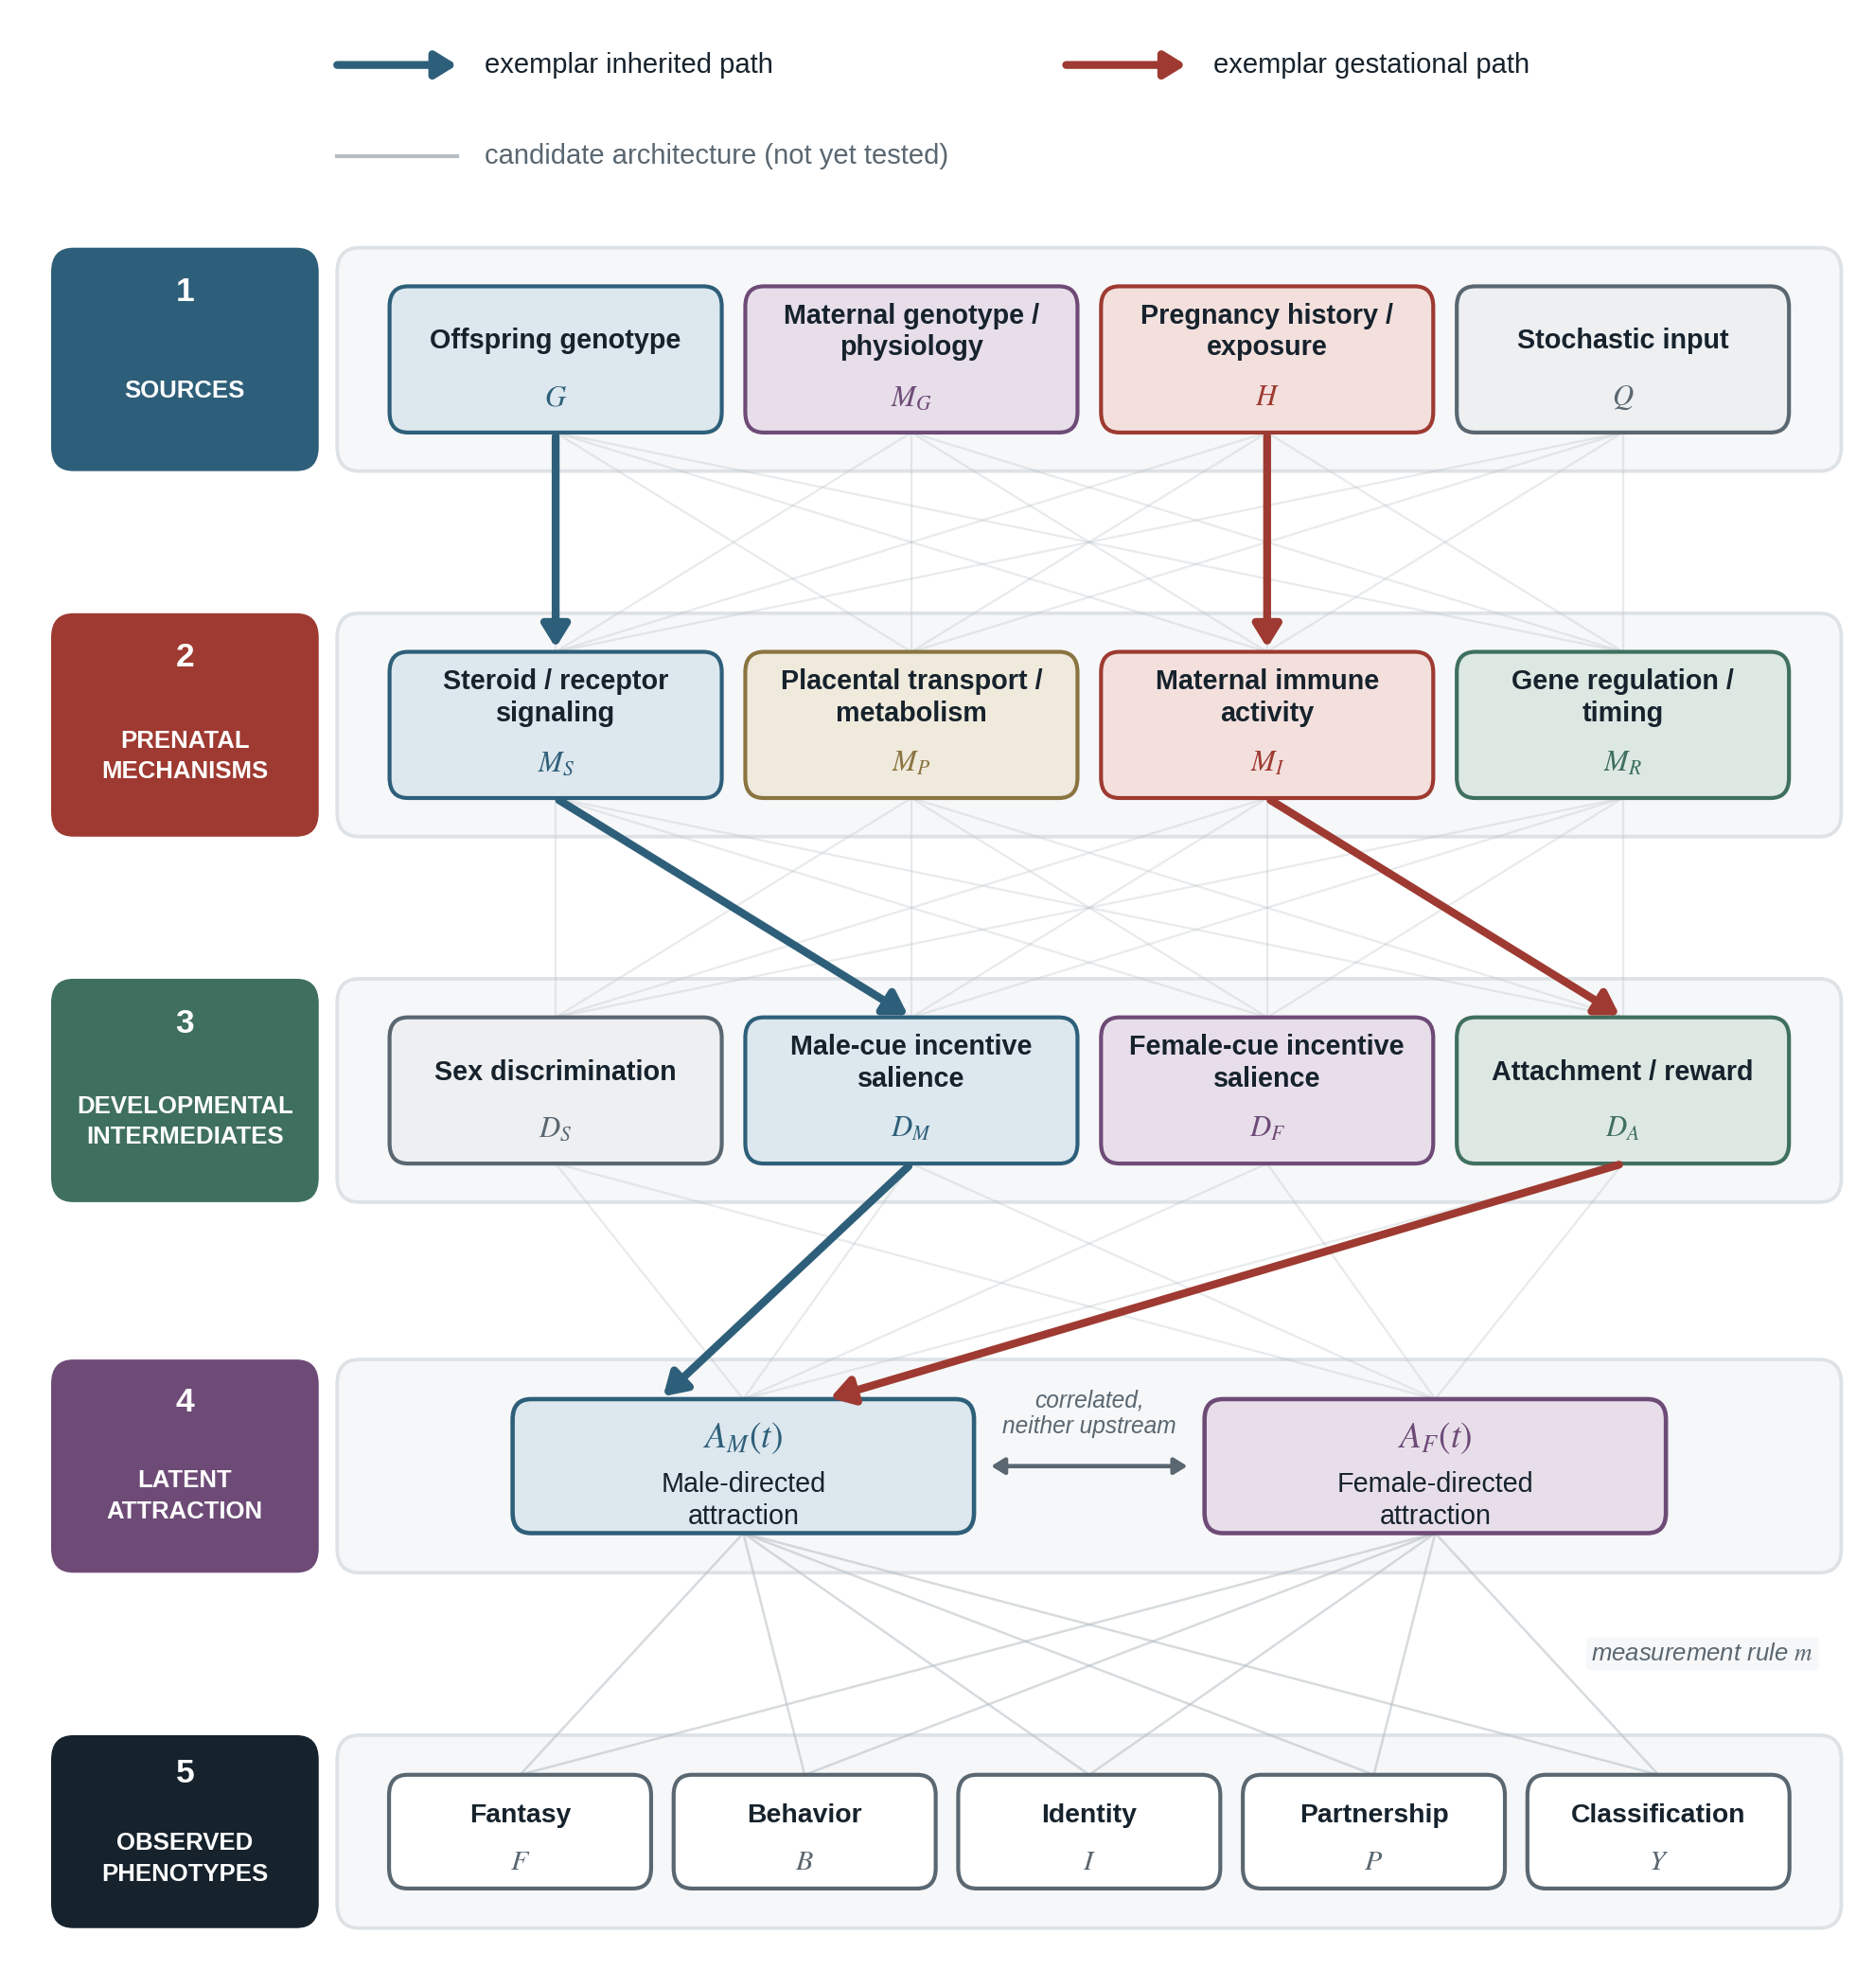
D3. Five causal levels are not collapsed. Offspring genotype, maternal genotype and physiology, pregnancy exposure, and stochastic variation are sources. Steroid signaling, placental transport, immune activity, and gene regulation are prenatal mechanisms. Sex discrimination, cue-specific incentive salience, attachment, and social reward are developmental intermediates. Repeated male-directed and female-directed attraction constitutes a distinct latent-attraction level. Fantasy, behavior, identity, partnership, and categorical classification are observed phenotypes or measurement rules. Diffuse polygenicity describes architecture, maternal-line concentration describes a transmission signature, and regulatory state describes a mechanism or modifier. None is automatically a route.
D4. Prevalence is a joint-distribution expectation, and all margins are stated on one scale. Cohort prevalence is the expectation of the outcome probability over inherited liabilities, maternal states, exposures, mechanisms, and their interactions. It is not the arithmetic sum of named route frequencies. This is descriptive standardization, not a theoretical result. Because the explanandum is latent attraction, every primary estimand and every equivalence margin is defined on the latent-attraction scale; quantities defined on the observed classification Y are derived from those through the fixed rule m, as set out below.
Empirical Propositions
P1. Male androphilia is a convergent phenotype produced by more than one mechanism-distinct prenatal path. The claim is stronger than generic multifactorial causation. At least two directed source-to-mechanism-to-intermediate paths must make material, replicable contributions to enduring attraction and must generate different temporal, conditional-independence, external-prediction, or intervention-relevant patterns. Material is numerical and is defined once, in Table 1, as a standardized mediated or identified edge-intervention effect on latent attraction of at least .10; a path qualifies only if it also shows a lower 90% confidence bound above .05 in an independent probability-weighted hold-out, a robustness value of at least .20, and support from at least one design that does not rely on sequential ignorability. Overlapping loadings and interactions are expected; sealed etiological classes are not.
P2. The inherited architecture is distributed, possibly omnigenic, and substantially shared across sexes, but its phenotypic channels need not be identical. No common high-penetrance locus is necessary or sufficient. Many small effects may enter several prenatal mechanisms and may be correlated through linkage disequilibrium, pleiotropy, or genome-wide regulatory coupling. The theory therefore does not require a small set of orthogonal inherited factors, and its evolutionary test partitions the full common-variant signal into mutually exclusive, preregistered bookkeeping bins that can each contain thousands of correlated effects.
P3. Prenatal organization is the principal causal stage, not a one-dimensional deficit in androgen exposure. Major biological effects on enduring attraction are established through prenatal or early perinatal organization of cue valuation and social reward. The proposition does not predict global under-masculinization, one direction of hormone effect in every subgroup, or a single anatomical signature. Steroid concentration, receptor sensitivity, placental metabolism, immune perturbation, cell-specific regulation, and developmental timing can generate different configurations.
P4. Evolutionary maintenance is component-specific and carries an explicit aggregate commitment. Inherited variation may persist through antagonistic pleiotropy, sexual antagonism, overdominance, mutation-selection balance, drift, linkage, or another quantitatively specified process, and different parts of the genome may be maintained by different processes. The inherited commitment is made at the level of the covered aggregate, not of any single favorable component: the compensatory-maintenance arm is tested with selection-valid, mutually exclusive full-genome partitions covering the prespecified majority of common-SNP heritability given in Table 1. Their carrier-weighted aggregate under natural fertility must be nonnegative after expression costs are included. The uncovered fraction is left unclassified: no effect estimated in the covered set is extrapolated to it, and it cannot rescue or condemn the covered portfolio. Pregnancy-dependent and stochastic paths do not require balancing selection on orientation, but the implicated vulnerability must be costly or constrained to remove.
Figure 1 maps the five causal levels and highlights two contrasting exemplar paths against the broader candidate graph.
Formal Causal Architecture
Sources, Mechanisms, and Paths
Let Si denote sources for male i: offspring inherited genotype, maternal genotype and physiology, pregnancy history and exposure, and stochastic developmental input. Let Mi denote prenatal mechanisms, Di developmental intermediates, and Ai = (AMi, AFi) the latent attraction state. Observed phenotypes Oi include fantasy, behavior, identity, partnership, and a categorical rule Yi. A minimal acyclic representation is:
Mi = fM(Si, UMi); Di = fD(Mi, Si, UDi); Ai = fA(Di, Mi, Si, UAi).
A route is a directed path from one element of Si through one or more elements of Mi and Di to Ai. Two routes are causally distinct only if the data support nonredundant temporal order, mediator dependence, conditional-independence restrictions, identified edge interventions, or external predictions. Mediation alone is insufficient because modest unmeasured mediator-outcome confounding can manufacture the same signature. A path therefore requires both confounding-robust prospective mediation and an independent identification source (the two-key rule). Different predictors loading on different factors is insufficient. Conversely, two causal routes may converge onto one smooth attraction distribution, so failure to recover latent classes does not demonstrate one etiology.
This graph resolves the causal-level problem. Diffuse polygenicity can feed steroid signaling, immune susceptibility, placental biology, neural gene regulation, or several simultaneously. Maternal-line concentration can arise from X linkage, maternal genotype, imprinting, interacting autosomal loci, correlated maternal exposure, or ascertainment. Maternal immune activity is a mechanism linked to pregnancy history. Regulatory state may mediate or modify any path. The directed graph, not the label, determines the route.
The Primary Phenotypes and the Classification Rule
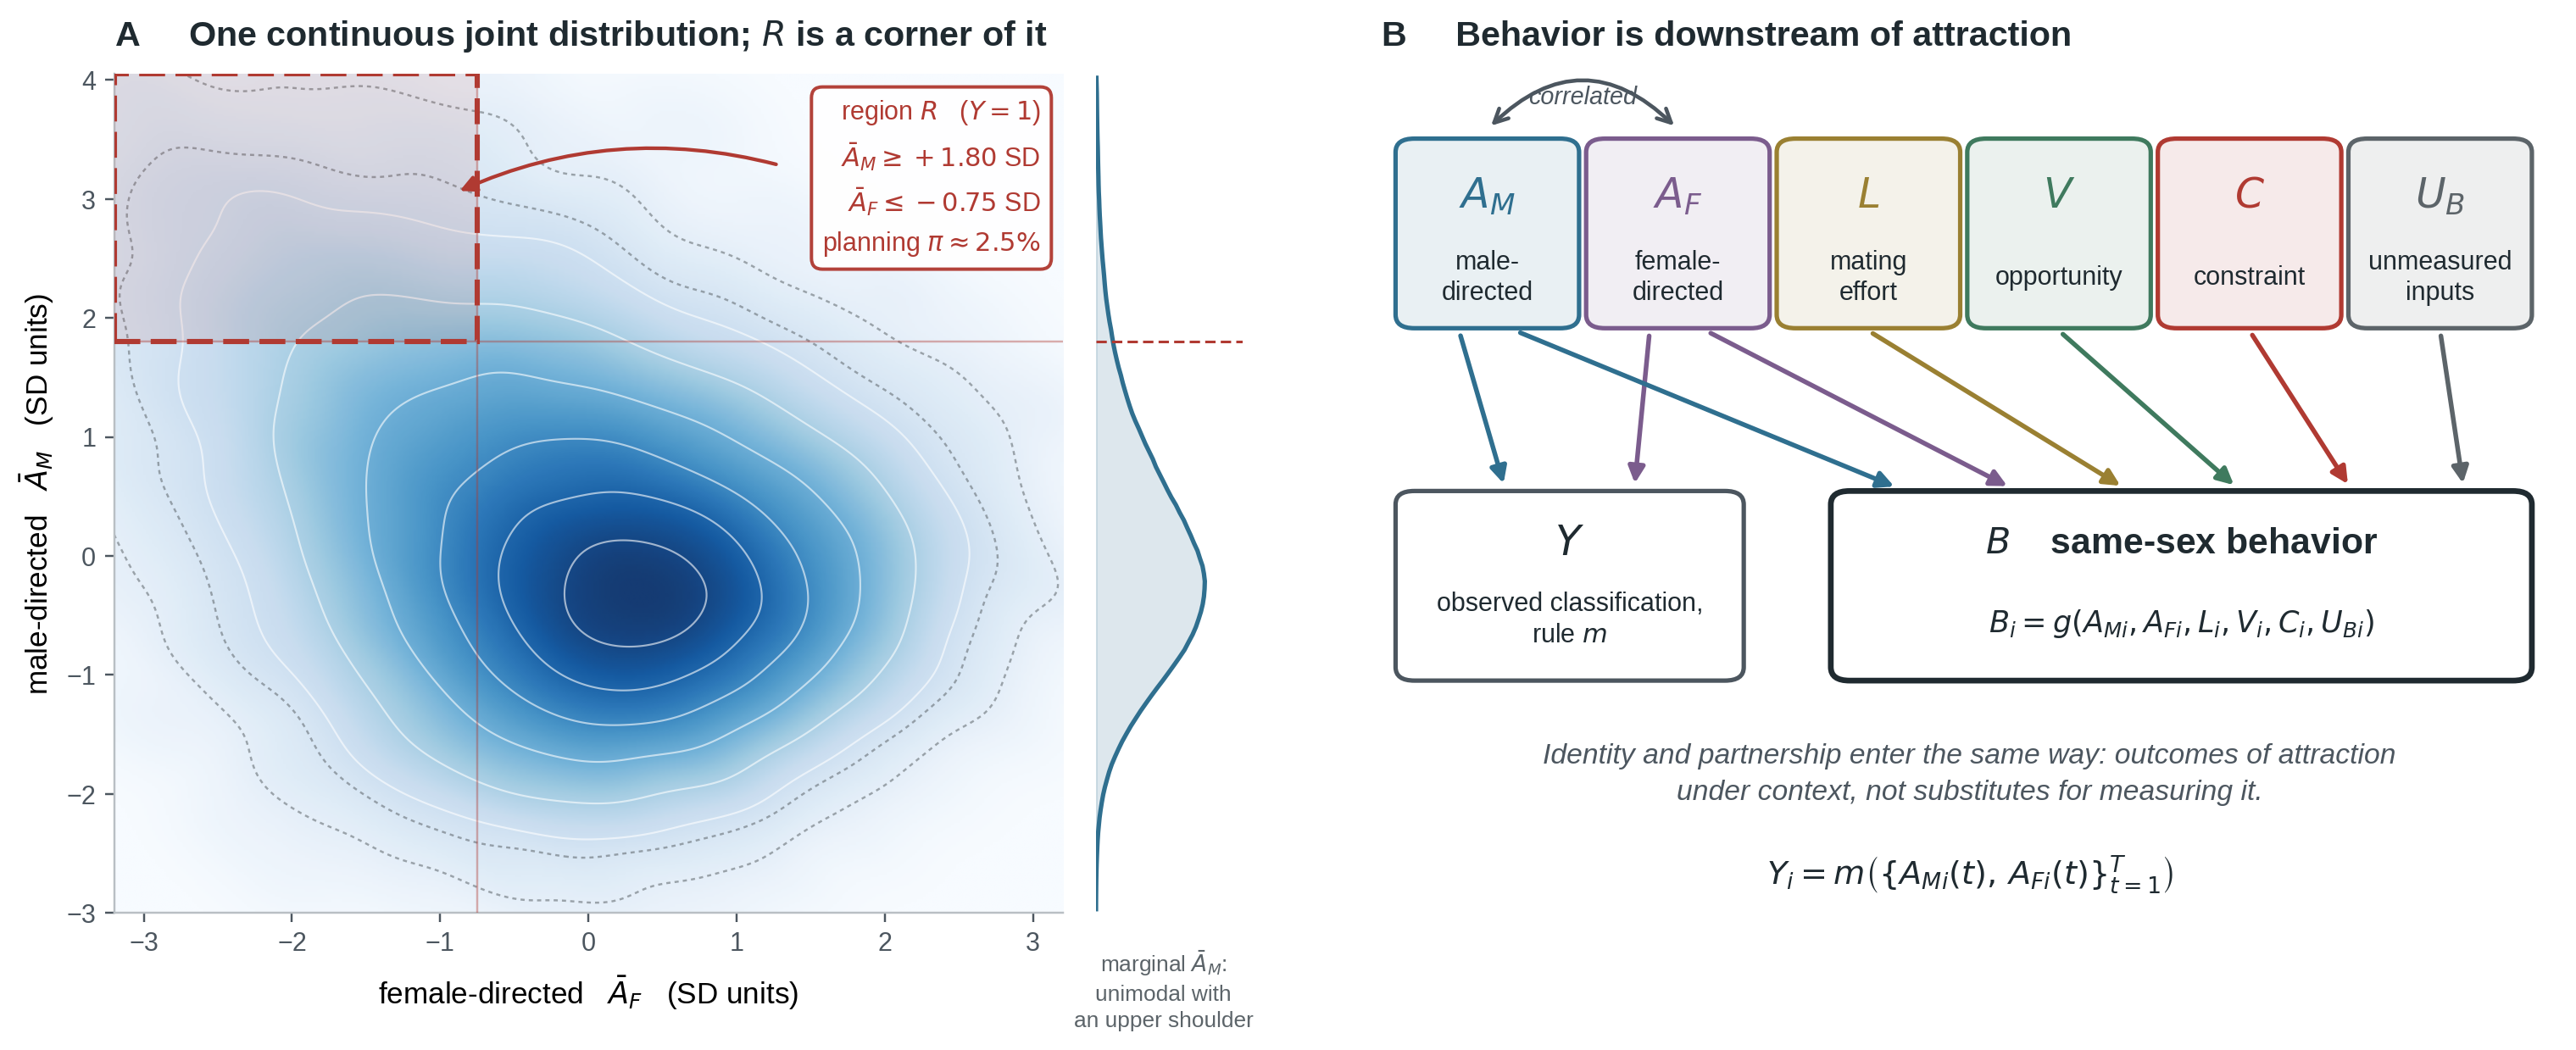
For each measurement occasion t, let AMi(t) denote latent male-directed attraction and AFi(t) latent female-directed attraction. Each is estimated from multiple indicators of intensity, physiological or subjective response, fantasy, persistence, and partner-specific incentive value. The primary adult quantities are the longitudinal summaries ĀMi and ĀFi, the person-level means across T ≥ 4 occasions spanning at least five years, standardized in the male population. They are not a one-dimensional Kinsey score and not an ever-versus-never behavior item.
The measurement rule m and the region R are specified here rather than named. The rule maps the repeated profile to its person-level means and applies the region:
Yi = m({AMi(t), AFi(t)} for t = 1, ..., T) = 1 if and only if ĀMi ≥ +1.80 and ĀFi ≤ −0.75.
Under the planning joint distribution described below, R contains approximately 2.5% of men, consistent with published estimates of predominant or exclusive androphilia in probability samples. The thresholds, the required number and span of occasions, and the standardizing population are fixed in advance and are reported in Table 1. The rule is a declared function of the latent summaries, not a separate construct: R is a corner of one continuous joint distribution, not a class recovered from the data.
Same-sex behavior is downstream. Let Li index general mating effort and behavioral propensity, Vi opportunity and partner availability, Ci social, legal, or interpersonal constraint, and UBi unmeasured behavioral inputs. Then behavior is represented as:
Bi = g(AMi, AFi, Li, Vi, Ci, UBi).
This removes the circularity in which partner-sex valuation was placed inside the function that defined orientation. Attraction is modeled directly; behavior is an outcome of attraction plus general propensity and context. A one-factor genomic result can therefore reject P2 without implying that attraction, behavior, identity, and partnership are interchangeable measurements.
Figure 2 shows the joint attraction distribution with the region R marked, and places the downstream behavioral and classification indicators.
Prevalence, Node Interventions, and Path-Specific Effects
Let Z contain measured sources, mechanisms, intermediates, covariates, and interactions, and let Ft be their joint distribution in cohort t. Population prevalence is:
πt = EZ ~ Ft[Pr(Y = 1 | Z)].
This expression is descriptive standardization over the cohort joint distribution. It is not a controlled-direct-effect formula, and the presence of intermediates inside Z does not make conditioning on them causally innocuous. The expression permits nonlinearities, overlap, and interaction. It also prevents the contradiction created by calling prevalence a simple sum of route incidences. Inclusion-exclusion is available only when routes are represented as binary sufficient-cause events. With continuous loadings and interaction, a case produced by moderate inherited liability plus moderate immune exposure has no unique natural assignment to either route.
A node intervention is a total effect of changing a source or mechanism, not a path-specific effect. For a prespecified node X and reference x0, define Δnode(X, t) = πt − πt{do(X = x0)}. Under this sign convention, a protective reference intervention that lowers prevalence yields a positive Δnode. This quantity inherits the standard assumptions of structural causal models and target-population transport (Hernán & Robins, 2020; Pearl, 2009).
For a prespecified set of directed edges P, the primary edge-intervention contrast is defined on latent attraction:
PSEP,t = Et[ĀM(x along P, x0 along P̄) − ĀM(x0)].
The corresponding contrast on Y is reported as a derived quantity obtained by applying m. The node, edge set, mediator order, target population, and treatment of interaction must be declared in advance. Edge-specific effects are not identified merely because all nodes are measured. A recanting witness or exposure-induced mediator-outcome confounder can block identification precisely when interacting paths share downstream nodes (Avin et al., 2005; Robins & Greenland, 1992; VanderWeele et al., 2014). When edge-identification conditions fail, report node or joint-intervention effects, interventional analogues, or bounds rather than path-attributable cases (Imai et al., 2010; VanderWeele, 2015). Shapley allocation is permissible only as a convention-dependent descriptive decomposition (Shapley, 1953).
The Robustness Gate Is a Curve, Not a Pass or Fail
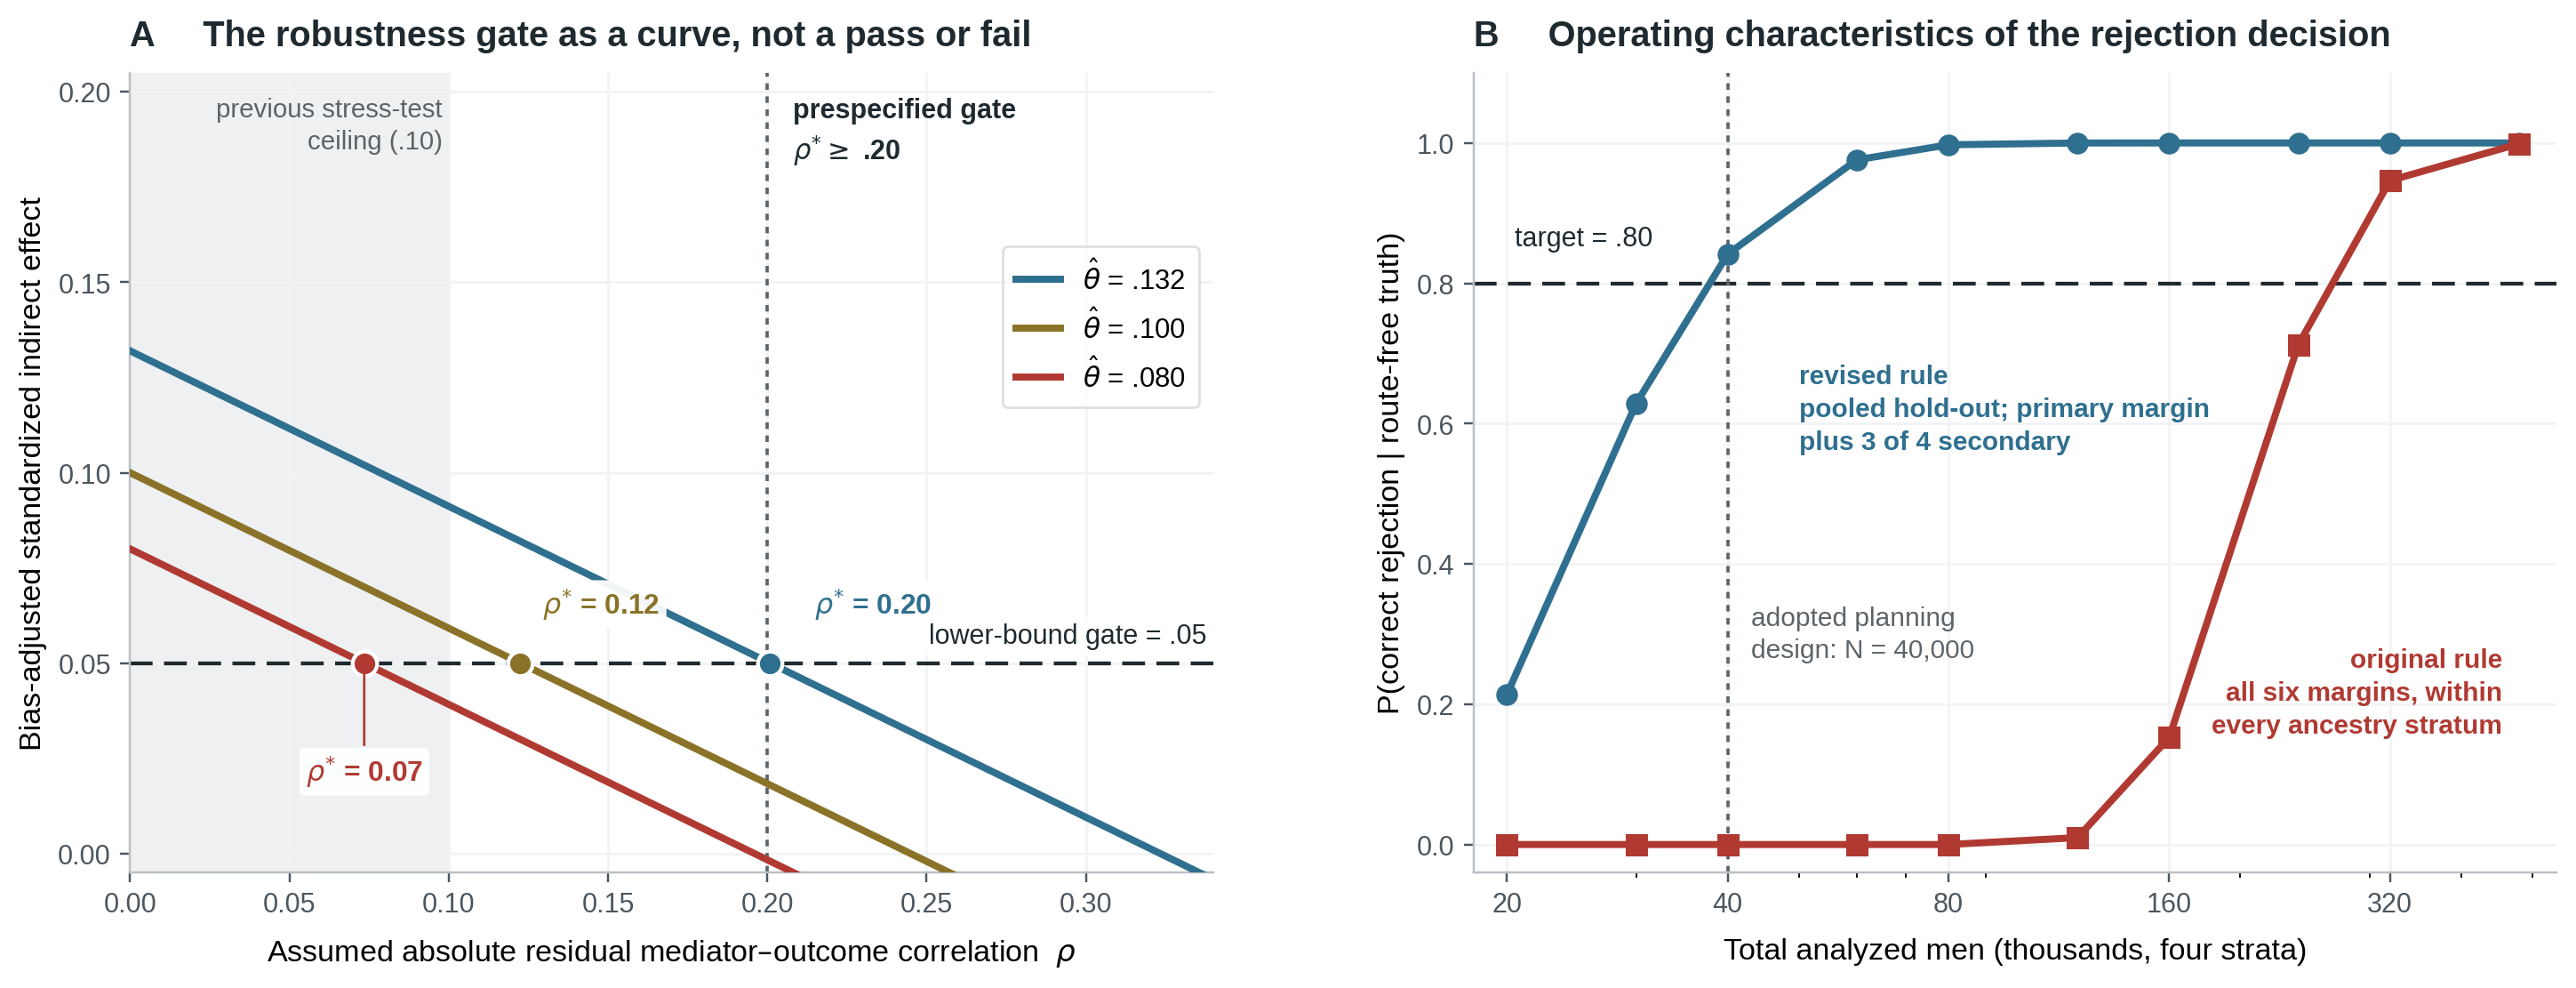
Every mediated-effect decision includes a prespecified residual-correlation sensitivity analysis. Rather than a single assumed ceiling, each candidate path is summarized by its robustness value ρ*, the smallest assumed absolute residual mediator-outcome correlation at which the path's lower 90% bound falls to .05 (Cinelli & Hazlett, 2020; VanderWeele & Ding, 2017). Under the linear model used in the worked example, the bias-adjusted standardized indirect effect is approximately θ̂ − aκρ, where a is the source-to-mechanism coefficient and κ is the ratio of the outcome to mediator residual standard deviations. The gate is ρ* ≥ .20.
Two consequences must be stated rather than left implicit. First, the materiality threshold and the robustness gate are not independent. With a = .40 and κ = 1.02, a path whose naive indirect effect is exactly .10 has ρ* = .12 (these planning values treat sampling error in the hold-out as negligible); requiring ρ* ≥ .20 therefore implies a naive indirect effect of about .132. The effective materiality threshold is a function of the gate, and both are reported. Second, a stress test fixed at an assumed residual correlation of .10, the ceiling adopted in an earlier version of this framework, is too lenient. The justification for .20 is given with the worked example below, and Figure 5A reports the full curve so that readers can apply their own ceiling instead of accepting a single pass-or-fail verdict.
Placing All Margins on One Scale
Materiality and the causal equivalence margin are defined on latent attraction A. Prevalence, Δnode, and the prevalence equivalence margin are defined on π = Pr(Y = 1), the output of the classification rule. These are not interchangeable scales, and a margin that looks strict on one can be permissive on the other. Because m and R are now fixed, the mapping can be computed rather than assumed.
For planning, the joint distribution of (ĀM, ĀF) is approximated as bivariate normal with correlation −.55. Under that approximation and the rule in Table 1, π = 2.53%, and the local derivative of prevalence with respect to a uniform shift δ in ĀM is dπ/dδ = .048. A standardized effect of .03 on latent attraction therefore corresponds to a prevalence difference of .0015, and the materiality threshold of .10 corresponds to .0048. Across plausible values of the attraction correlation (−.45 to −.65) and threshold pairs yielding π between 1.7% and 3.6%, the derivative ranges from .036 to .058 and the derived margin from .0011 to .0017.
This computation shows why the prevalence margin cannot be fixed independently. A prevalence equivalence margin of .005, which looks strict and was the value fixed in an earlier version of this framework, corresponds to .103 standardized units on latent attraction, approximately the materiality threshold itself and about 3.4 times the causal margin of .03. Two models differing by a full material effect would have satisfied it. The prevalence margin is therefore derived as the image of the .03 causal margin under m, equal to .0015 under the planning assumptions and recomputed from the fitted joint distribution in the analysis. The remaining predictive margins are secondary, are reported on their own conventional scales, and never carry the primary decision.
This also resolves an apparent tension with D1. The explanandum is latent attraction; Y is a declared deterministic function of the latent summaries; quantities on Y are therefore derived reports of the same underlying causal structure rather than a second, categorical explanandum.
The Decision Rule and Its Operating Characteristics
The causal core is rejected when both of the following hold in the pooled, probability-weighted hold-out: fewer than two paths pass the materiality-and-identification gate, and the route-free comparator is statistically equivalent to the best path model on the primary causal margin and on at least three of the four secondary predictive margins. Equivalence is assessed with two one-sided tests at 90% confidence (Lakens et al., 2018). The route-free comparator is defined as a structural causal model with exactly one or two latent continuous liabilities, no source-specific mediator map, and no identified edge effects; three- to five-liability versions are mandatory sensitivity models. The core is retained only when at least two paths pass the gate and the comparator fails equivalence; every other pattern, such as a single qualifying path without comparator equivalence, is reported as indeterminate rather than as retention.
Three features of this rule depart from the more obvious design, a conjunctive rule that requires equivalence on every margin within every stratum, which an earlier version of this framework used and which cannot fire at any feasible sample size.
First, the primary decision is made in the pooled hold-out rather than separately within every genetic-similarity stratum. Requiring equivalence within every stratum makes the least-powered stratum the binding constraint and gives the perverse result that adding strata makes the theory harder to reject. Stratum-specific estimates are now reported as heterogeneity diagnostics. If stratum-specific point estimates for the primary contrast differ by more than the causal margin, the pooled verdict is withheld and the result is reported as heterogeneous rather than as retention.
Second, equivalence is required on the primary causal margin and a prespecified majority of the secondary margins rather than on every margin conjunctively. Establishing equivalence is harder than failing to reject a difference, because a two one-sided test requires the whole interval to fall inside the margin. Requiring roughly two dozen such conclusions, every margin in each of four strata, guarantees indeterminacy rather than falsification.
Third, the operating characteristics of the rejection decision are reported alongside those of retention. This is the substantive point. Under the conjunctive rule, with 5,000 analyzed men per stratum and a 30% hold-out, the half-width of the 90% interval for the primary contrast is 1.645 × .026 = .042, which exceeds the .03 margin, so no dataset of that size can declare equivalence at all. Simulation across a design grid confirms the consequence: the probability of a correct rejection under a route-free data-generating process was below .001 for every design up to 80,000 analyzed men and did not reach .80 until roughly 250,000. Under the pooled rule the same probability is .21 at 20,000, .84 at 40,000, and .98 at 60,000 (Figure 5B). The planning design is therefore 10,000 analyzed men in each of four strata, a total of 40,000, with a 30% hold-out.
The final sample is the smallest value in a preregistered grid for which end-to-end Monte Carlo recovery of the complete decision procedure reaches at least .90 probability of a correct retention under a portfolio generator and at least .80 probability of a correct rejection under a route-free generator, with familywise false-decision probability at or below .05. The simulation must include the case oversample, design weights, hold-outs, stratum diagnostics, missingness, nonnormality, measurement error, interaction, ascertainment, and the mediator-outcome confounding scenarios used in the robustness gate.
Feasibility follows from the prevalence and can now be derived rather than asserted. Enrolling 10,000 men in the region R, a 25% oversample of the analyzed sample, requires screening approximately 395,000 eligible responders at π = 2.5%. At a 20% screening response and 50% enrollment, that is roughly 4.0 million invitations; at a 25% response and 60% enrollment, roughly 2.6 million. A design with a smaller oversample of 5,000 requires between 1.3 and 2.0 million. These figures are large, and they are stated rather than hidden.
A note on what is claimed. The theory is falsifiable by prespecified adverse patterns whose operating characteristics are stated above. Critical Studies 1 through 4 and 6 are runnable with current technology at the sample sizes given. Critical Study 5 is not, because no existing resource combines a genome-wide study of repeated attraction with directly genotyped natural-fertility outcomes; it is a long-horizon falsifier. Describing the whole program as decisively falsifiable would overstate what is presently available, and the distinction is maintained throughout.
Competing Models and Discriminating Predictions
The causal-portfolio theory extends, but does not originate, the proposition that similar adult orientations may arise through multiple biological processes. Rahman and Wilson (2003) synthesized genetic, prenatal endocrine, neurodevelopmental, and evolutionary evidence without a path-identification framework. VanderLaan and colleagues later argued explicitly for biodevelopmental subgrouping and for using gender expression and other markers to carve orientation at its joints (VanderLaan et al., 2022). The present theory is distinct only if it earns the additional commitments listed below: routes must be directed paths through a causal graph, phenotype production must be separated from observation, node effects must not be mislabeled as path effects, and each inherited or recurrent component must be paired with its own evolutionary accounting rule.
Table 2 states the closest alternatives and applies one decision logic throughout. The theory is not supported merely because several predictors are significant, several factors fit, or several mechanisms are biologically plausible. It survives only when at least two paths pass the materiality-and-identification gate and the joint pattern rejects the alternatives.
| Model | Causal claim | Discriminating prediction | Decision pattern |
|---|---|---|---|
| Causal-portfolio theory | At least two mechanism-distinct prenatal paths contribute materially; paths may overlap and share downstream nodes. | Each path clears the materiality-and-identification gate in Table 1; inherited and recurrent components require different accounting. | Retained only if at least two paths pass that gate and route-free comparators fail equivalence under the pooled rule. |
| Route-free continuous liability | Several continuous dimensions influence one developmental outcome, without separable causal paths. | Dimensions improve fit, but no material, confounding-robust, independently identified source-specific path remains. | Favored when one or two liabilities are equivalent to the best path model on the causal margin and a majority of secondary margins. |
| Biodevelopmental profile heterogeneity | Biomarker-defined developmental profiles or subgroups contribute to similar orientations. | Subgroups or profiles replicate, but evolutionary maintenance and counterfactual prevalence effects need not be specified. | Favored if replicated profiles do not yield additional causal, prevalence, or maintenance predictions. |
| Broad polygenic architecture plus measurement | One distributed inherited architecture produces variation; apparent routes reflect phenotype definition, ascertainment, or proxy error. | Direct mechanism measures add no material, confounding-robust prediction beyond polygenic liability and reliable attraction measures. | Favored if endocrine, immune, placental, and regulatory indicators produce no qualifying path. |
| Independent genetic and gestational mechanisms | Genetic and maternal mechanisms both exist but require no integrated portfolio or interaction accounting. | Main effects replicate, but overlap, interaction, and joint-distribution predictions are unnecessary. | Favored if the integrated model provides no additional calibrated prediction or counterfactual utility. |
| Mutation-selection without compensation | Inherited liability is recurrently introduced and selected against; no carrier advantage is required. | Covered inherited components show negative selection under natural fertility, with allele frequencies explained by mutation, drift, or linkage. | Favored for the maintenance arm if covered inherited components remain negative and mutation, drift, or linkage explains their frequencies. |
Note. The same decision logic is applied to every row. The theory is not supported because several predictors are significant, several factors fit, or several mechanisms are biologically plausible.
A Worked Example: Same Data, Different Identification Assumptions
Two simulations are reported. They answer different questions and their numbers must not be pooled. Simulation A fits competing structural models to one portfolio-generated dataset in which there is no true residual mediator-outcome correlation; it asks whether explicit causal restrictions can defeat a route-free competitor. Simulation B sets a true residual mediator-outcome correlation and asks how badly a naive mediation analysis is degraded by it. Table 3 reports Simulation A; the confounding-degradation results are given in the text and in Figure 5A.
| Model or quantity | Causal restrictions | Value |
|---|---|---|
| Strict portfolio SCM | G → MG → A and I → MI → A; no direct source-to-attraction paths; no residual M–A correlations | −9.870 |
| Strict route-free SCM | G and I predict A directly; MG and MI have no remaining link to A | −9.922 |
| Route-free sensitivity SCM | Direct source effects plus freely estimated residual MG–A and MI–A associations | −9.870 |
| Strict-portfolio advantage over the strict route-free model | Difference in held-out log likelihood per observation | +.052 |
| Fitted residual correlations in the sensitivity model | ρ(eMG, eA); ρ(eMI, eA) | .225; .236 |
Note. SCM = structural causal model. n = 20,000; seed 20260825; 70% training and 30% hold-out; four attraction indicators with reliability .70; true standardized indirect effect .10 for each path; no true residual mediator-outcome correlation. The first three values are held-out Gaussian log likelihoods per observation; the fourth is a difference in those units; the fifth is a pair of correlations. Simulation B, in which a true residual mediator-outcome correlation was imposed, is reported in the text and in Figure 5A.
Simulation A: Model Comparison on One Dataset
With n = 20,000 and seed 20260825, independent standardized sources G and I each caused its mechanism with coefficient .40, and each mechanism caused latent attraction A with coefficient .25. The true standardized indirect effect for each path was therefore .10, exactly the materiality threshold. Four attraction indicators had reliability .70. Models were estimated in 70% of the sample and evaluated in the same 30% hold-out.
The strict portfolio structural causal model achieved a held-out Gaussian log likelihood of −9.870 per observation. A strict route-free model, in which G and I acted directly on A and the mechanism markers had no remaining link to A, fit worse at −9.922. That difference of .052 log-likelihood units per observation shows that a genuinely route-free model can be rejected when its causal restrictions are made explicit. But a route-free sensitivity model that allowed residual MG–A and MI–A associations fit at −9.870, essentially tying the portfolio model, with fitted residual correlations of .225 and .236.
Those two fitted values require careful interpretation, and the interpretation determines the robustness gate. They are not estimates of the residual confounding present in the data, because there was none: they arise because a model deprived of the mediator paths must absorb genuine mediation as residual covariance. What they do establish is that residual correlations of roughly .23 are observationally admissible in this design: that is the confounding a skeptic needs in order to explain away a true path of exactly the materiality threshold, and the data cannot exclude it. A stress test capped at an assumed residual correlation of .10 therefore never reaches the region in which the competing explanation lives. No gate below .23 can make a single mediation estimate immune to that skeptic, and a gate at .23 or above would require naive effects of about .144 and so exclude true effects at the threshold itself. The gate is set at ρ* ≥ .20 so that a path must survive most of the admissible range, and the exposure that remains is closed by the second key, independent identification, rather than by a stricter ceiling.
Simulation B: Confounding Degradation
In a separate simulation the structural coefficients remained .40 and .25, so each causal indirect effect remained .10, but a true residual mediator-outcome correlation was imposed. At a true residual correlation of .10, a naive no-confounding mediation analysis estimated the two indirect effects at approximately .136 and .136. At .20, the corresponding estimates were approximately .167 and .168. These are not small distortions around the decision boundary.
The example changes the burden of proof. A strict route-free model can misfit the same data that generated a material path, but observational equivalence returns as soon as plausible residual mediator-outcome association is admitted. The causal-portfolio theory therefore requires the two-key rule in Table 1: confounding-robust prospective mediation plus an identification source that does not rely on sequential ignorability. Factor solutions, marginal associations, and mediation regressions alone cannot establish a route. Figure 5A reports the robustness curve for three candidate effect sizes, and Figure 5B the operating characteristics of the rejection decision.
Empirical Constraints on the Causal Graph
Inherited Sources: Polygenic Liability and Maternal-Line Signatures
Twin, family, linkage, and association studies establish familial and inherited influence without high penetrance. Concordance and heritability estimates vary with phenotype, recruitment, prevalence assumptions, nonresponse, and model specification. That variation is compatible with causal heterogeneity but is not evidence for it by itself. The invariant conclusion is narrower: inherited liability is distributed, probabilistic, and accompanied by large individual-specific variation (Bailey & Pillard, 1991; Bailey et al., 1999, 2000; Kendler et al., 2000; Långström et al., 2010).
Genome-wide results support highly distributed effects and permit an omnigenic architecture in which many variants influence attraction through broadly expressed regulatory networks (Boyle et al., 2017). Broad same-sex behavior shares genetic signal with attraction, fantasy, and identity, but the measures are not equivalent; the same GWAS also showed differences between ever-versus-never behavior and proportion of same-sex partners (Ganna et al., 2019, 2021; Hamer et al., 2021). Reported common-SNP heritability for same-sex sexual behavior ranged from approximately .08 to .25 depending on phenotype and sample; the planning value adopted for repeated attraction is .15, and the actual denominator for the coverage criterion must be estimated in the discovery sample rather than assumed. Functional or genomic partitions can summarize common-SNP heritability without pretending that each partition is a sparse causal factor (Finucane et al., 2015). The theory predicts substantial shared variance between attraction and behavioral propensity while exposing their residual separation to direct genomic test.
Xq28 is a contested source signature, not a route. Selected brother-pair studies reported excess sharing, an independent Canadian study did not support a large effect, unselected family histories did not show the expected general maternal-line excess, and later linkage scans produced ascertainment-sensitive results (Bailey et al., 1999; Hamer et al., 1993; S. Hu et al., 1995; Mustanski et al., 2005; G. Rice et al., 1999; Sanders et al., 2015). Extreme maternal X-inactivation skewing and the Han Chinese associations at FMR1NB and ZNF536 remain biologically relevant leads, but neither establishes one maternally inherited path (Bocklandt et al., 2006; S.-H. Hu et al., 2021). A valid inherited path requires a replicated molecular signal, a mechanism linking it to prenatal development, and predicted family and fitness consequences in held-out samples.
Prenatal Endocrine, Placental, and Regulatory Mechanisms
The organizational-activational principle supplies the strongest general mechanism for the sexual differentiation of partner preference. Early steroid action alters gene expression and neural organization, whereas adult hormones primarily regulate libido and activation rather than reverse the target of attraction (Hines, 2011; Phoenix et al., 1959). Swift-Gallant et al. (2023) substantially strengthen this part of the theory. Across experiments of nature and converging biomarkers, the review concludes that early gonadal hormone action influences human sexual orientation, with particularly strong evidence that androgens increase gynephilia in people raised as girls and suggestive evidence that estrogens increase androphilia. That evidence does not by itself establish the direction or magnitude of ordinary variation among typically developing males.
The same review also prevents an overstatement. Evidence for within-male variation is mixed: men with typical external anatomy may have passed a threshold sufficient for genital masculinization while differing in tissue-specific sensitivity, developmental timing, immune exposure, or nonlinear hormone effects. Swift-Gallant et al. explicitly consider multiple male pathways and the possibility that both relatively low and relatively high androgenic conditions could be associated with different nonheterosexual subgroups. The causal-portfolio theory therefore locates the bottleneck at prenatal organization but rejects a scalar deficit model.
Placental transport and metabolism are causal mechanisms in their own right because fetal tissue exposure is not identical to maternal serum concentration. Receptor expression, local steroid conversion, co-regulator availability, fetal genotype, maternal genotype, and cell-specific regulatory state can redirect or buffer the same circulating signal. Regulatory variation is not an unexplained residual class. It is a set of measurable mechanisms that must predict tissue-relevant transcription, interact with upstream exposures, and survive prospective and functional replication.
Canalization is used in its conventional sense of developmental robustness. Thresholding, monozygotic discordance, epigenetic persistence, and stochastic divergence can occur within a robust system, but none demonstrates canalization by itself. The transgenerational epigenetic model of W. R. Rice et al. (2012) remains a candidate implementation rather than a required premise. Adult blood or saliva methylation classifiers (Ngun et al., 2015) cannot reconstruct a prenatal neural state, and reprogrammed cells undergo extensive epigenetic resetting while retaining memory of the adult source tissue rather than of any prenatal neural state (Kim et al., 2010). A regulatory claim becomes causal only when prospective prenatal measures, identified mediation, and perturbation converge.
Maternal Immune and Pregnancy-History Mechanisms
The fraternal birth-order association is the strongest evidence for a pregnancy-dependent influence not reducible to the son's inherited orientation liability. Many studies report higher odds of male homosexuality with additional older biological brothers, and the maternal immune hypothesis proposes progressive immunization to male-specific fetal proteins (Blanchard, 2004, 2018; Blanchard & Bogaert, 1996; Blanchard & Klassen, 1997; Bogaert, 2006). Bogaert et al. (2018, 2022) reported the predicted pattern for selected NLGN4Y sequences, but the study was cross-sectional, used maternal samples decades after pregnancy, and required a sequence-specific correction. It is a mechanism-linked lead, not a demonstrated mediator.
Older-brother count is an exposure proxy rather than a route indicator. Population-attributable fractions near 15% and 29% depend on different exposure definitions and assumptions and do not assign individual cases (Blanchard & Bogaert, 2004; Cantor et al., 2002). The current logistic approach of Blanchard et al. (2025) separates older-brother, older-sister, and sibship-size contrasts under explicit ceteris-paribus substitutions and recommends theory-neutral labels for the resulting coefficients. Dutch register results found birth-order effects not confined to men, and simulations show that some family-size controls can induce or exaggerate an association (Ablaza et al., 2022; Vilsmeier et al., 2023). The immune path therefore requires serial within-woman immunization, index-pregnancy fetal exposure, prospective prediction of adult attraction, and mediation of a material part of the biological-older-brother association across a preregistered antigen and cellular panel.
Developmental Intermediates, Behavior, and Comparative Theory
The graph includes sex discrimination because comparative work shows that same-sex sexual behavior can arise from mating decisions without same-sex attraction. N. W. Bailey and Zuk (2009) established the general evolutionary distinction, and Lerch (2026) connects it to classic mate-choice theory: imperfect sex discrimination, acceptance thresholds, encounter rates, opportunity costs, mating effort, and social context can generate or maintain same-sex behavior. Lerch explicitly distinguishes same-sex attraction from behavior and does not claim that the animal framework directly explains human sexuality. Its importance here is causal: it supplies a competing path from general mating strategy to behavior that bypasses enduring male-directed valuation.
This distinction is also visible within the human genomic record. Risk-taking can mediate a positive fertility association for bisexual behavior while exclusive same-sex behavior remains negatively associated with reproduction (Song & Zhang, 2023, 2024). A behavioral-propensity component may therefore have a different evolutionary history from the component that produces high ĀM and low ĀF. Critical Study 2 asks whether that difference survives reliable attraction measurement and multivariate genomic modeling; it does not assume the answer.
Comparative evidence sets boundaries rather than supplying a direct human explanation. Same-sex behavior is phylogenetically widespread, plastic, and sometimes socially functional. Comparative reconstructions support repeated gains, losses, and associations with sociality rather than one assumed ancestral state (Gómez et al., 2023; Monk et al., 2019). In male rhesus macaques it was common, repeatable, modestly heritable, and not detectably associated with reduced reproduction in one intensively studied population (Clive et al., 2023). Male-oriented rams are more directly relevant because a stable minority chooses males even when estrous females are available, demonstrating that enduring male-oriented partner preference can arise in a mammal without human identity or culture (Roselli et al., 2011). None of these findings establishes homology with human androphilia.
Neurobiological Convergence Without a Unitary Brain Type
The expected common endpoint is functional, not gross anatomical. Interacting hypothalamic, striatal, oxytocin, vasopressin, dopamine, attachment, and social-reward systems are plausible substrates of cue valuation and pair bonding (Blumenthal & Young, 2023). Different upstream paths can converge on relative incentive value without producing one homosexual brain type.
The postmortem and imaging record is too inconsistent to define a route. INAH3 is sexually dimorphic, but the best-controlled replication did not confirm an orientation difference in volume or neuron number; the anterior commissure and suprachiasmatic findings remain unreplicated or were replaced by different molecular observations; and HIV/AIDS ascertainment confounded most homosexual postmortem samples (Allen & Gorski, 1992; Byne et al., 2001; S.-H. Hu et al., 2021; LeVay, 1991; Swaab & Hofman, 1990). A systematic review found conflicting results, extensive overlap, small samples, and no consistent phenotype (Frigerio et al., 2021). A larger UK Biobank imaging-genetics analysis detected small multivariate cross-sex shifts, predominantly in women, and regional associations with a same-sex-behavior polygenic score; its broad behavioral phenotype and modest effects do not identify a prenatal path (Abé et al., 2021). Route-stratified neurobiology becomes informative only after upstream causal measures define the strata.
Evolutionary Accounting
Inherited Components
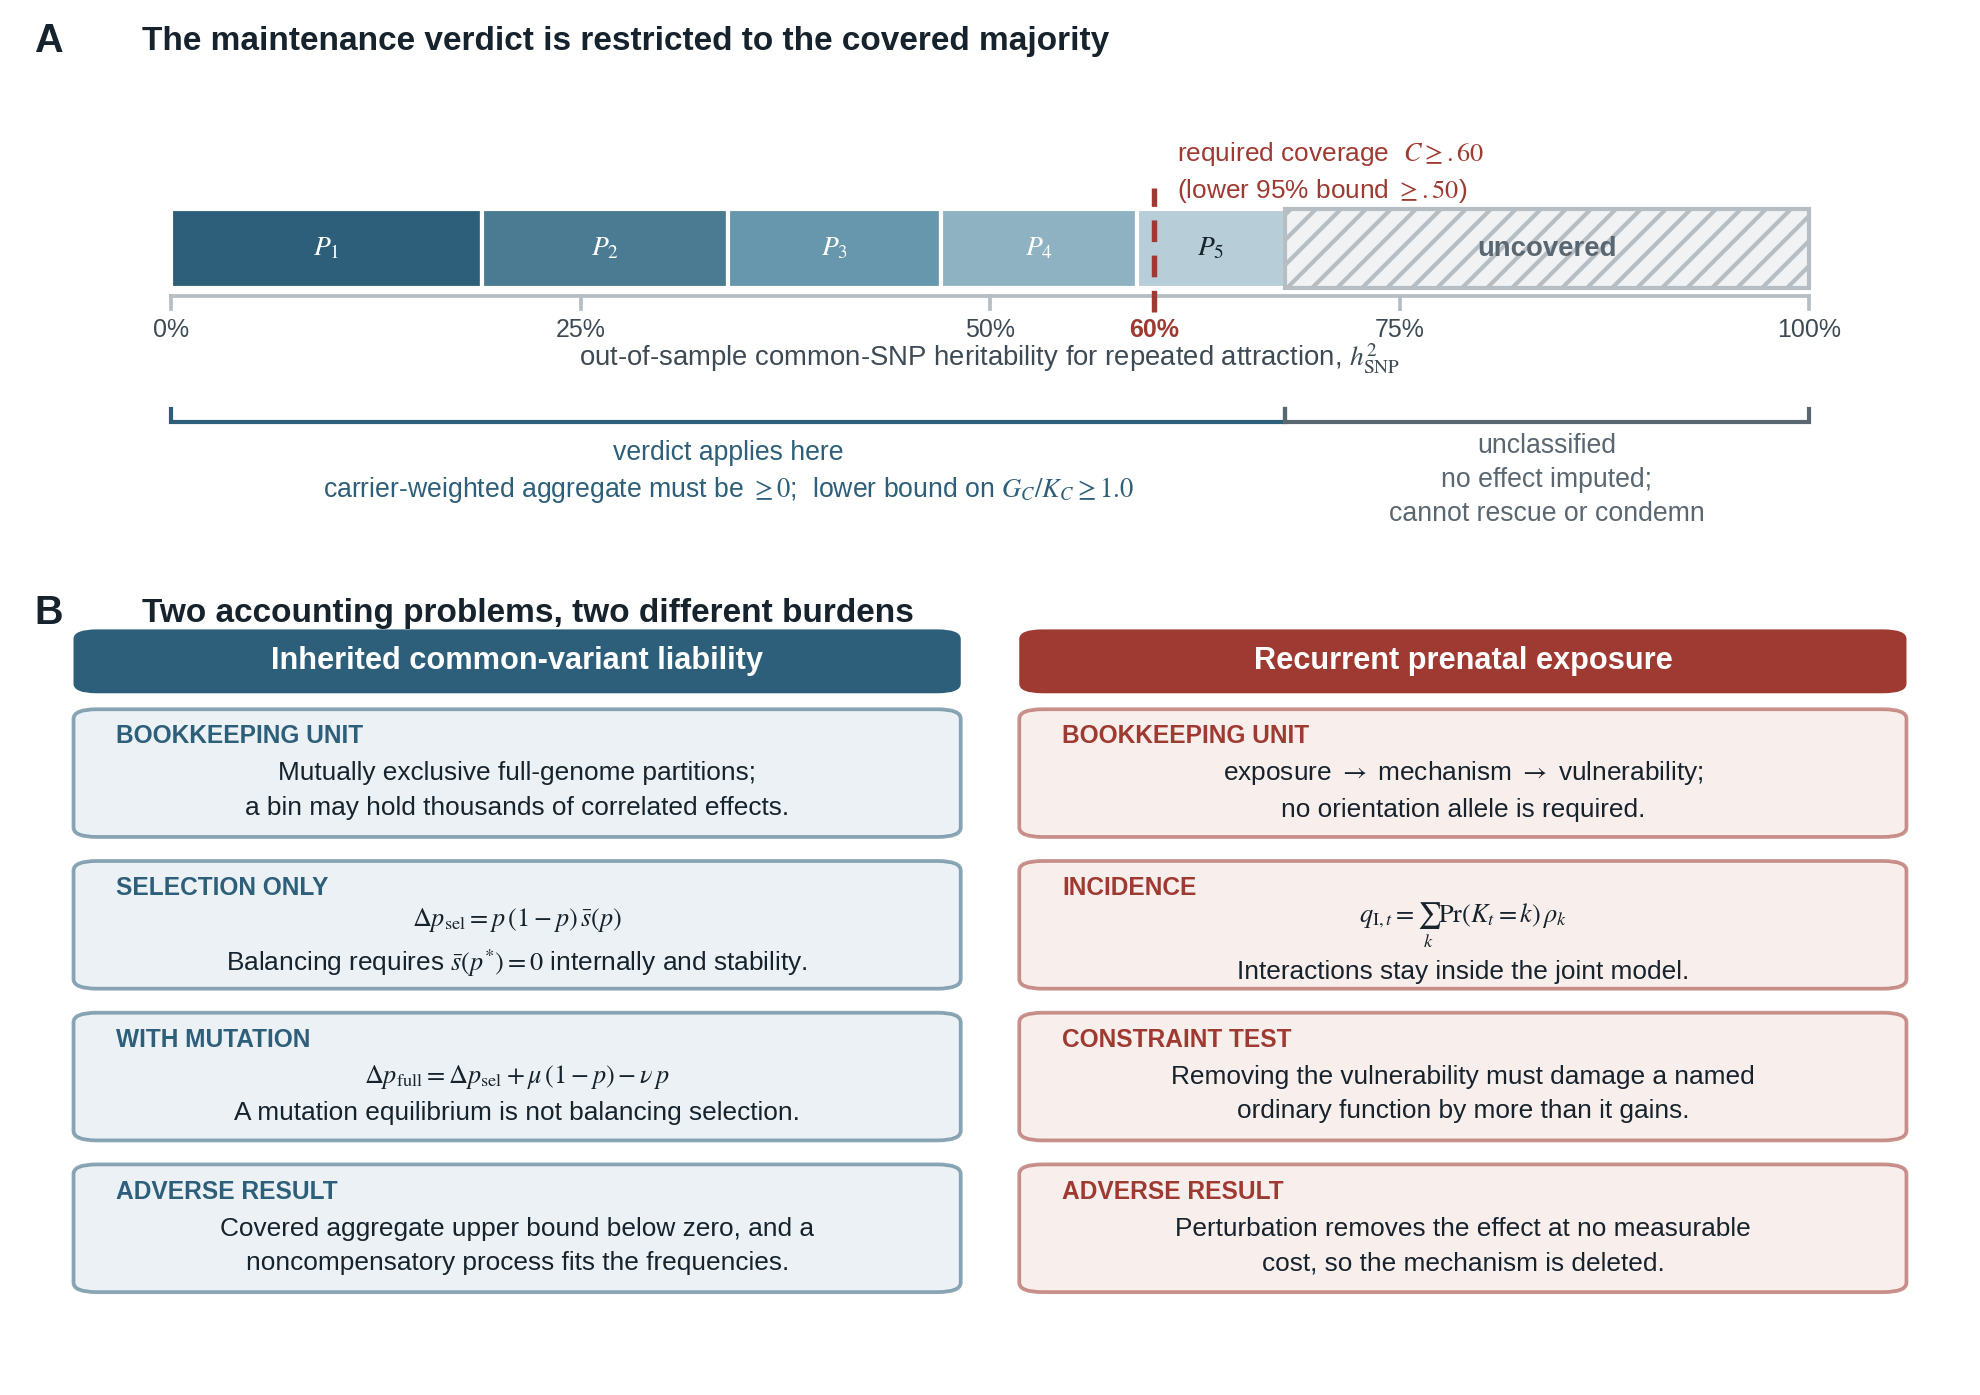
For inherited allele or molecular component Aj with frequency pj,t, separate selection from mutation. The selection-only change is Δpsel = pj(1 − pj) s̄j(pj,t). Balancing selection is admissible only when s̄j(pj*) = 0 at an internal frequency and the selection-only recurrence is locally stable. With reversible mutation, Δpfull = Δpsel + μj(1 − pj) − νj pj. If s̄j is negative at every frequency, mutation can still create an internal equilibrium, but that equilibrium is mutation-selection balance, not balancing selection. Antagonistic pleiotropy, sexual antagonism, and overdominance predict different dominance, chromosomal, frequency, and sex-specific signatures (Gavrilets & Rice, 2006).
The carrier ledger includes heterosexual men, androphilic men, women with different attraction profiles, and every other phenotype through which the component alters reproduction. Positive effects on partner number, paternal care, mate appeal, or female fertility matter only if their carrier frequencies and effect sizes offset the cost in expressed androphilic males. Miller's (2000) threshold arithmetic shows how a small advantage in numerous nonexpressing carriers could offset a large cost in a rare high-liability configuration under its illustrative allele-frequency assumptions, but the arithmetic is not evidence that the required coupling exists.
Current evidence is mixed. Heterosexual men with nonheterosexual twins reported more opposite-sex partners, and broad same-sex-behavior signal correlates genetically with opposite-sex partner number (Zietsch et al., 2008, 2021). Risk-taking explains the advantageous association for bisexual behavior in contemporary data (Song & Zhang, 2024). The desirable-dad hypothesis predicts a pathway through paternal care and mate value but lacks direct natural-fertility confirmation (Felesina & Zietsch, 2024). Modern partner number is not historical reproductive success, and a broad behavior score is not a score for enduring androphilia.
The Explicit Covered-Portfolio Commitment
The compensatory-maintenance claim is deliberately stronger than an existential search for one favorable component. Let the full common-variant signal be divided into mutually exclusive, preregistered genome partitions: functional annotations, nonoverlapping linkage-disequilibrium blocks, or disjoint score bins. A partition may contain thousands of correlated effects and is not assumed to be an orthogonal biological factor. A partition is selection-valid only after out-of-sample heritability or score replication, within-family validation, measurement invariance, ancestry-specific linkage-disequilibrium calibration, and transport to a natural-fertility regime. Let C be the fraction of out-of-sample SNP heritability covered by selection-valid partitions. The coverage and compensation criteria are given in Table 1.
For the covered partitions, let KC be the expected reproductive cost attributable to expression in androphilic males under natural fertility and GC the carrier-weighted gain in all other male and female contexts. No selection effect is imputed to the uncovered fraction 1 − C. Sensitivity analyses may display hypothetical residual effects across an explicitly labeled range, but those assumptions do not enter the verdict and cannot turn an extrapolation into a bound.
Modern polygenic scores cannot simply be carried into historical populations. Critical Study 5 requires ancestry- and cohort-specific linkage-disequilibrium calibration, phenotype measurement invariance, within-family validation, direct genotyping in the earliest available natural-fertility cohorts, explicit gene-by-environment and fertility-regime interaction, and modeling of assortative mating. Ancestral imputation is secondary sensitivity analysis. Feasibility is stated rather than hidden: no existing dataset combines a genome-wide study of repeated attraction with all of these gates and directly genotyped natural-fertility outcomes. Planning simulations must therefore examine discovery samples of 250,000, 500,000, and 1,000,000 or more participants, followed by natural-fertility validation resources, and select the lowest design that attains the coverage and precision criteria.
The adverse covered-portfolio pattern is exact. The compensatory-maintenance claim for the covered majority fails when the coverage and transport criteria are met, the one-sided upper bound on aggregate selection across the covered partitions remains below zero, and a fitted mutation-selection, drift, or linkage process reproduces their frequencies without compensation; the noncompensatory account then replaces compensation for that majority (Table 2). If the covered aggregate is negative but no noncompensatory process reproduces the observed frequencies, the covered ledger is reported as unbalanced: compensation is not demonstrated, and the discrepancy between measured selection and observed frequency is not read as evidence of unmeasured compensation. Developmental heterogeneity could survive either result. Compensation for the selection-valid covered majority would not.
Recurrent Prenatal Exposures and Developmental Constraint
A pregnancy-history-dependent immune event can recur in every generation without orientation being vertically inherited. Its incidence depends on the distribution of prior male gestations and the conditional probability that a specified immune process contributes after each history. This solves an incidence question, not the evolutionary question of why the causal immune specificity, placental transfer, fetal target, or downstream sensitivity persists.
The theory therefore requires a node-specific trade-off. A modification that blocks the causal vulnerability must reduce expected fitness through impaired immunity, placental function, ordinary neural differentiation, or another identified process by more than it gains through preventing the rare outcome. Generic statements that immunity or developmental robustness is useful are inadmissible. If targeted perturbation removes the relevant effect without a measurable cost, the constraint claim fails and that mechanism is deleted.
Maternal-Line Fecundity, Kin Investment, and Social Effects
Early pedigree studies reported higher fecundity among mothers and maternal female relatives, and multilocus sexually antagonistic models showed that such a pattern could maintain a low-frequency liability (Camperio Ciani & Pellizzari, 2012; Camperio Ciani et al., 2004, 2008; Iemmola & Camperio Ciani, 2009). The pooled evidence now weighs against a general female-fecundity effect: common-method analyses, a decade of pooled Samoan data, Dutch registers, Czech data, and unselected pedigrees have been unfavorable (Ablaza et al., 2022; Bailey et al., 1999; Blanchard et al., 2020; Fořt et al., 2024; Semenyna et al., 2023). A rare marker-defined sex-antagonistic component remains testable, but it is not an established pillar and disappears if marker-selected families show no predicted female or heterosexual-carrier benefit.
Kin investment is subsidiary. Samoan fa'afafine report elevated avuncular willingness, whereas a Western community study found no general family-directed resource transfer and greater estrangement on some measures (Bobrow & Bailey, 2001; Vasey & VanderLaan, 2010). The inclusive-fitness requirement is quantitative: actual transfers must raise recipient survival and reproduction enough to offset forgone direct reproduction. Questionnaire differences, coalition correlations, and altered mating markets do not satisfy that criterion. They become maintenance mechanisms only when a genetic pathway to reproductive effects is demonstrated.
Figure 3 places the inherited and recurrent accounting problems side by side and states the coverage, covered-portfolio, and constraint gates.
Critical Tests: How the Theory Can Fail
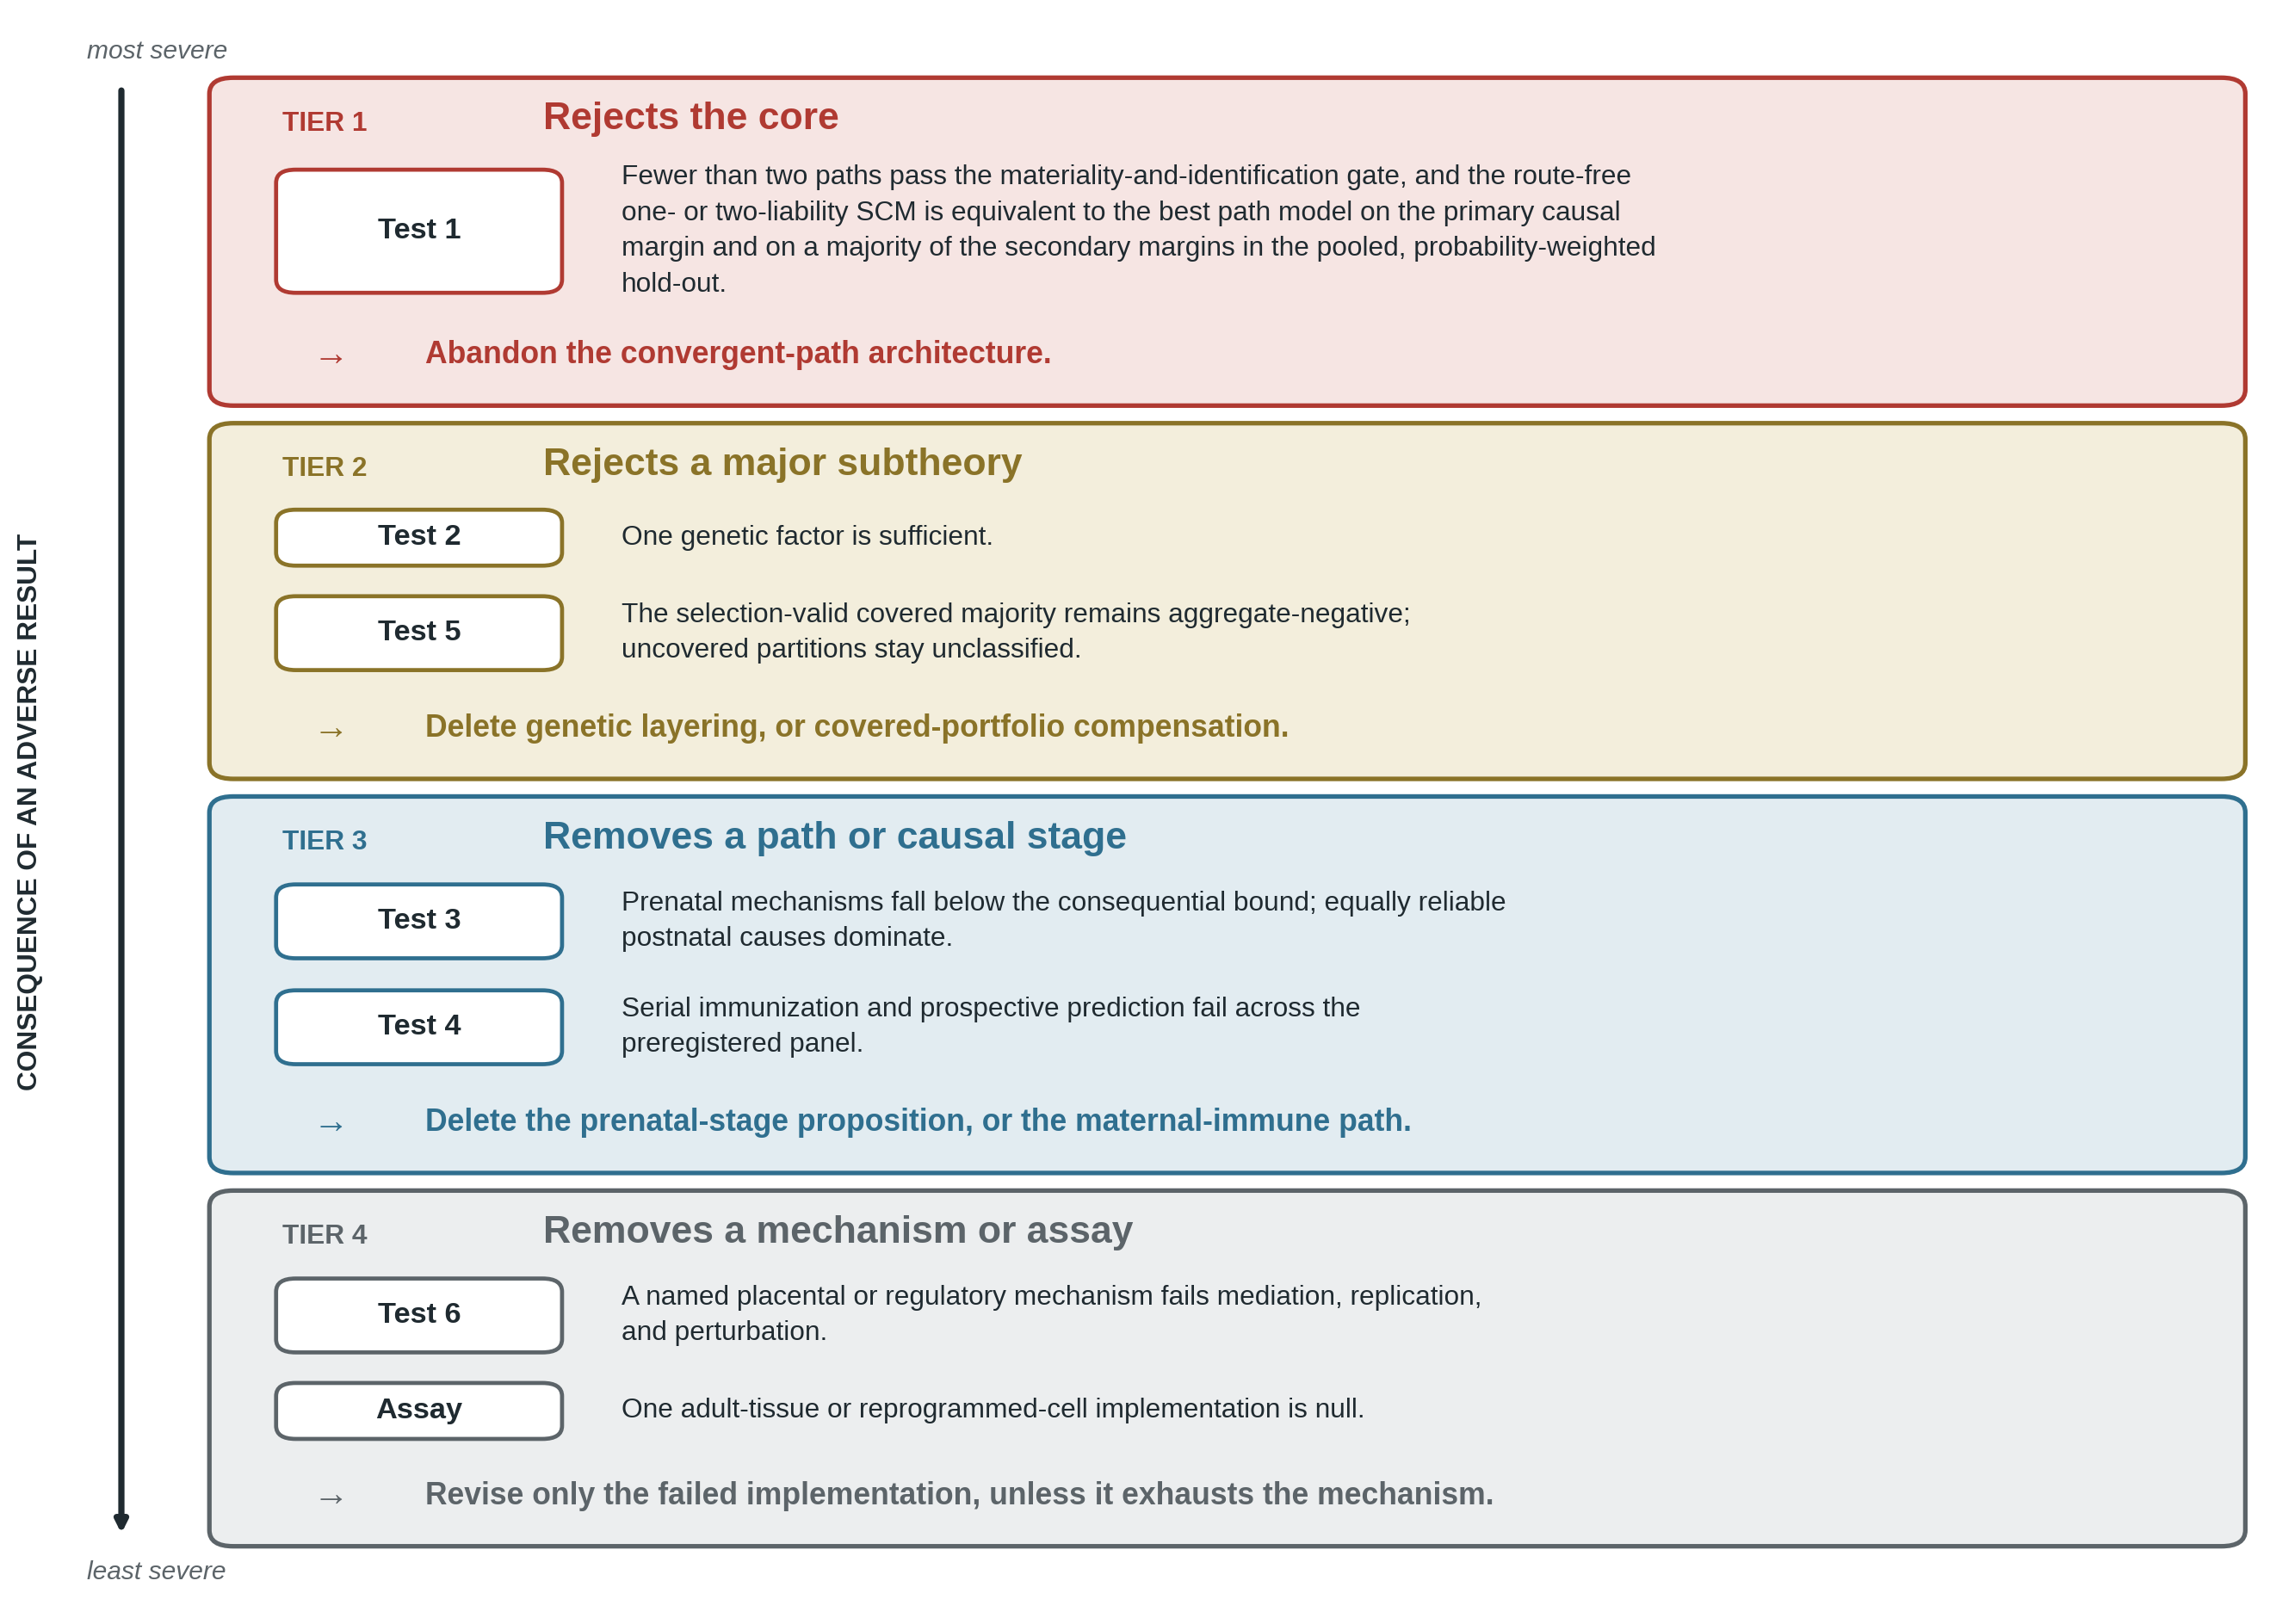
The critical studies are not a collection of desirable future projects. Each places a stated claim at risk and defines what follows from a negative result. Full sampling plans, the end-to-end simulation requirement, measurement requirements, and adverse decision rules appear in the Supplementary Material. All thresholds and margins referenced below are those in Table 1. The hierarchy is deliberately asymmetric: one result rejects the causal-portfolio core, two reject major subtheories, others delete a candidate path or mechanism, and assay-specific nulls remain assay-specific.
Critical Study 1: Do Mechanism-Distinct Causal Paths Exist?
Claims at risk: P1, with D3 and D4 as supporting commitments. Use probability-based screening in four genetic-similarity strata and retain inclusion probabilities for every screened and enrolled participant. Oversample the region R for precision, but carry inverse-probability or calibration weights into structural, predictive, and prevalence analyses. Begin the preregistered grid at 10,000 analyzed men per stratum, the planning design derived above, and adjust in either direction as the end-to-end simulation requires. A candidate path passes only if it clears the materiality-and-identification gate in Table 1. The core is rejected only under the pooled decision rule stated above, with stratum-specific results reported as heterogeneity diagnostics.
Critical Study 2: Are Attraction and General Behavioral Propensity Genetically Distinguishable?
Claim at risk: P2. Conduct a biobank-scale multivariate genomic study with reliable independent measures of ĀM, ĀF, fantasy, identity, behavior, partner proportion, sociosexuality, risk-taking, openness, mating effort, and reproductive outcomes in both sexes. Compare one-factor and layered genomic structural models with measurement error and cross-sex covariance modeled explicitly. The genetic-layer claim fails if one factor is decisively preferred, attraction and behavior are predictively equivalent in held-out samples, and no valuation-specific heritable component survives the prespecified equivalence bound. That result does not make orientation, behavior, identity, and partnership the same phenotype; it removes the proposed two-layer genetic architecture.
Critical Study 3: Is Prenatal Organization the Principal Causal Stage?
Claim at risk: P3. Follow a prospective pregnancy cohort with fixed gestational measurement windows for steroid concentrations, receptor sensitivity, placental transport and metabolism, immune activity, fetal genotype, and repeated developmental outcomes through adult attraction. The primary analysis compares causal models in which prenatal measures mediate inherited and maternal effects against models in which equally reliable postnatal variables dominate after temporal and familial confounding are addressed. The consequential bound is numbered rather than named: the proposition fails if every adequately measured prenatal mechanism has a standardized effect below .10 with a lower 90% bound below .05 in every powered path stratum, while postnatal predictors produce robust, temporally ordered, out-of-sample effects that clear the same bound. An average null is insufficient when opposing nonlinear path effects were not measured or powered.
Critical Study 4: Does a Pregnancy-History-Dependent Maternal Immune Path Exist?
Claim at risk: the named gestational immune path. Enroll women from first pregnancy, collect serial samples in fixed windows of every subsequent pregnancy, assay corrected NLGN4Y sequences and a preregistered broad Y-antigen and cellular panel, measure placental transfer and fetal exposure, and follow sons to repeated ĀM and ĀF measures. The specified NLGN4Y path fails if its serial and prospective predictions fail. The broader immune path is deleted if no male-gestation-specific immunization occurs across the preregistered panel, index-pregnancy exposure does not predict attraction, and the mechanism does not mediate a material fraction of the older-brother association within precise bounds.
Critical Study 5: Was the Covered Inherited Portfolio Nonnegative Under Natural Fertility?
Claim at risk: P4. Apply the components from Critical Study 2 only after the coverage, portability, ancestry, and measurement criteria in Table 1 are met. Use directly genotyped natural-fertility cohorts, validated pedigrees, within-family analyses, both sexes, expressed and nonexpressed carriers, completed reproduction, survival of descendants, assortative mating, and fertility-regime interaction. Estimate KC, GC, GC/KC, partition-specific and aggregate covered selection, and allele-frequency predictions. The compensatory claim for the covered majority is rejected when its one-sided aggregate upper bound is below zero and a fitted noncompensatory process explains the observed frequencies; a negative aggregate that no fitted process explains is reported as an unbalanced ledger, not as compensation. Uncovered partitions remain unclassified; a nonportable score or modern partner-count association cannot alter the verdict.
Critical Study 6: Does a Named Placental or Regulatory Mechanism Carry a Causal Path?
Claim at risk: a specified placental or regulatory implementation. Combine prospective placental, cord, and neonatal measures with genetically informed designs and targeted perturbation in tissue-relevant cellular or animal systems. The named mechanism must temporally predict attraction-relevant development, mediate an upstream source effect, interact as predicted with another path when interaction is claimed, and change a relevant developmental endpoint when perturbed. It is deleted if all four requirements fail within prespecified bounds. A null adult blood classifier or a null after induced-pluripotent reprogramming rejects only that assay because neither reconstructs prenatal neural regulation.
Figure 4 separates rejection of the core from rejection of genetic layering, covered-portfolio compensation, a prenatal path, or one assay.
Evolutionary Synthesis
The theory makes one decisive causal declaration: enduring male androphilia is not merely a high score on one or two route-free liabilities. It is a convergent attraction phenotype to which at least two mechanism-distinct prenatal paths make material contributions, in the precise sense given in Table 1. Those paths may share inherited sources, interact within the same pregnancy, and converge onto the same smooth attraction distribution. They are distinguished by causal restrictions and identified interventions, not by circles, clusters, factor labels, or a mediation regression alone.
The theory makes a second decisive evolutionary declaration: no single maintenance process carries the entire phenotype. Inherited common-variant liability is tested through selection-valid full-genome partitions and allele-level accounting across both sexes. The verdict is confined to the covered majority; uncovered partitions are not assigned the effect distribution of the covered set. Recurrent prenatal exposure faces an incidence and constraint problem. Observed prevalence is generated by their joint distribution. A candidate path that lacks its predicted robust causal evidence, compensation, or constraint is deleted rather than absorbed into an unspecified remainder.
Recent work sharpens rather than dilutes the theory. Swift-Gallant et al. (2023) strengthen prenatal organization while showing why male variation cannot be reduced to a single monotonic androgen scale. Lerch (2026) shows, through classic mate-choice theory, how same-sex behavior can emerge from sex discrimination, acceptance thresholds, and mating strategy without same-sex attraction. Together these findings require the architecture to separate prenatal mechanisms from developmental intermediates and attraction from behavior.
The framework is therefore more than an umbrella only insofar as its additional predictions survive. It must outperform multidimensional continuous liability, VanderLaan-style profile heterogeneity, broad polygenicity plus measurement, independent mechanisms without integrated accounting, and mutation-selection explanations without carrier benefit. The comparison, the indeterminate regions, and the outcomes that would defeat the theory are all explicit, and mediation alone is explicitly denied the power to establish a route.
Conclusion
Male androphilia is a minority endpoint of sexually differentiated development whose evolutionary persistence must be explained at the level of its causal components. Offspring genotype, maternal genotype and physiology, pregnancy exposure, and stochastic variation act through prenatal endocrine, placental, immune, and regulatory mechanisms. These mechanisms alter sex discrimination, cue-specific incentive salience, attachment, and social reward, producing repeated male-directed and female-directed attraction. Behavior, identity, and partnership follow from that attraction under additional motivational and social constraints; they are not substitutes for it.
The conventional paradox is not dissolved by saying that causes differ. It is replaced by a stricter set of questions. Which directed paths are causal? Which effects survive explicit mediator-outcome confounding sensitivity and independent identification? Which edge-specific effects are identified, and which require joint interventions or bounds? Which selection-valid covered partitions were nonnegative under natural fertility? Which recurrent vulnerabilities are constrained or costly to remove?
The causal-portfolio theory answers with five noncollapsed levels, a numbered materiality threshold, a two-key path-identification rule, a robustness gate reported as a curve, an explicit distinction between node and path effects, a single scale on which every margin is stated, a majority-coverage criterion without extrapolation to the uncovered fraction, and adverse results that compel abandonment or deletion. Fewer than two qualifying paths plus route-free equivalence under the pooled decision rule rejects the core. A negative selection-valid covered portfolio rejects compensatory maintenance for that majority. Failure of a named immune, endocrine, placental, or regulatory mechanism removes that path.
Two limits are stated plainly rather than absorbed. The rejection decision has adequate operating characteristics only at sample sizes near 40,000 analyzed men, and the corresponding recruitment burden runs to millions of invitations. Critical Study 5 cannot presently be run at all. The theory is falsifiable in the specific sense that its adverse patterns and their error rates are prespecified, not in the looser sense that a decisive study is currently available. Nothing here is protected by the word heterogeneity.
Declarations
Funding. No funding was received to assist with the preparation of this manuscript. No funds, grants, or other support were received for conducting this work.
Competing interests. The author has no competing interests to declare that are relevant to the content of this article. The author operates MooreMetrics, the independent website on which this manuscript is published; it is otherwise unrelated to the subject matter of the article.
Ethics approval. Not applicable. This is a theoretical article. It reports no new data collected from human participants or animals. All empirical findings discussed are drawn from the published literature, and all quantitative results reported by the author are derived from simulated data.
Availability of data and material. No new empirical data were generated. The simulated datasets underlying the worked example, the robustness curve, and the decision-rule operating characteristics are fully specified by the generating parameters and random seeds reported in the article and can be reproduced from them.
Code availability. The simulation and figure-generation scripts are available from the author on request.
Use of generative AI and AI-assisted technologies. The author used Claude Opus 4.8, GPT-5.6 Sol Pro, and Grok 4.3 to prepare initial drafts and for proofreading. The author reviewed and edited all content and takes full responsibility for it.
References
Abé, C., Lebedev, A., Zhang, R., Jonsson, L., Bergen, S. E., Ingvar, M., Landén, M., & Rahman, Q. (2021). Cross-sex shifts in two brain imaging phenotypes and their relation to polygenic scores for same-sex sexual behavior: A study of 18,645 individuals from the UK Biobank. Human Brain Mapping, 42, 2292–2304. https://doi.org/10.1002/hbm.25370
Ablaza, C., Kabátek, J., & Perales, F. (2022). Are sibship characteristics predictive of same sex marriage? An examination of fraternal birth order and female fecundity effects in population-level administrative data from the Netherlands. The Journal of Sex Research, 59, 671–683. https://doi.org/10.1080/00224499.2021.1974330
Allen, L. S., & Gorski, R. A. (1992). Sexual orientation and the size of the anterior commissure in the human brain. Proceedings of the National Academy of Sciences of the United States of America, 89, 7199–7202. https://doi.org/10.1073/pnas.89.15.7199
Apostolou, M. (2022). The direct reproductive cost of same-sex attraction: Evidence from two nationally representative U.S. samples. Archives of Sexual Behavior, 51, 1857–1864.
Avin, C., Shpitser, I., & Pearl, J. (2005). Identifiability of path-specific effects. In Proceedings of the 19th International Joint Conference on Artificial Intelligence (pp. 357–363).
Bailey, J. M., & Pillard, R. C. (1991). A genetic study of male sexual orientation. Archives of General Psychiatry, 48, 1089–1096.
Bailey, J. M., Dunne, M. P., & Martin, N. G. (2000). Genetic and environmental influences on sexual orientation and its correlates in an Australian twin sample. Journal of Personality and Social Psychology, 78, 524–536. https://doi.org/10.1037/0022-3514.78.3.524
Bailey, J. M., Pillard, R. C., Dawood, K., Miller, M. B., Farrer, L. A., Trivedi, S., & Murphy, R. L. (1999). A family history study of male sexual orientation using three independent samples. Behavior Genetics, 29, 79–86.
Bailey, N. W., & Zuk, M. (2009). Same-sex sexual behavior and evolution. Trends in Ecology & Evolution, 24, 439–446. https://doi.org/10.1016/j.tree.2009.03.014
Blanchard, R. (2004). Quantitative and theoretical analyses of the relation between older brothers and homosexuality in men. Journal of Theoretical Biology, 230, 173–187.
Blanchard, R. (2018). Fraternal birth order, family size, and male homosexuality: Meta-analysis of studies spanning 25 years. Archives of Sexual Behavior, 47, 1–15.
Blanchard, R., & Bogaert, A. F. (1996). Homosexuality in men and number of older brothers. American Journal of Psychiatry, 153, 27–31.
Blanchard, R., & Bogaert, A. F. (2004). Proportion of homosexual men who owe their sexual orientation to fraternal birth order: An estimate based on two national probability samples. American Journal of Human Biology, 16, 151–157. https://doi.org/10.1002/ajhb.20006
Blanchard, R., & Klassen, P. (1997). H-Y antigen and homosexuality in men. Journal of Theoretical Biology, 185, 373–378.
Blanchard, R., Kabátek, J., & Zdaniuk, B. (2025). A current approach to logistic regression analysis of birth order and sexual orientation. Archives of Sexual Behavior, 54, 4013–4022. https://doi.org/10.1007/s10508-025-03275-3
Blanchard, R., Krupp, J., VanderLaan, D. P., Vasey, P. L., & Zucker, K. J. (2020). A method yielding comparable estimates of the fraternal birth order and female fecundity effects in male homosexuality. Proceedings of the Royal Society B: Biological Sciences, 287, Article 20192907. https://doi.org/10.1098/rspb.2019.2907
Blumenthal, S. A., & Young, L. J. (2023). The neurobiology of love and pair bonding from human and animal perspectives. Biology, 12, Article 844.
Bobrow, D., & Bailey, J. M. (2001). Is male homosexuality maintained via kin selection? Evolution and Human Behavior, 22, 361–368. https://doi.org/10.1016/S1090-5138(01)00074-5
Bocklandt, S., Horvath, S., Vilain, E., & Hamer, D. H. (2006). Extreme skewing of X chromosome inactivation in mothers of homosexual men. Human Genetics, 118, 691–694. https://doi.org/10.1007/s00439-005-0119-4
Bogaert, A. F. (2006). Biological versus nonbiological older brothers and men's sexual orientation. Proceedings of the National Academy of Sciences of the United States of America, 103, 10771–10774.
Bogaert, A. F., Skorska, M. N., Wang, C., Gabrie, J., MacNeil, A. J., Hoffarth, M. R., VanderLaan, D. P., Zucker, K. J., & Blanchard, R. (2018). Male homosexuality and maternal immune responsivity to the Y-linked protein NLGN4Y. Proceedings of the National Academy of Sciences of the United States of America, 115, 302–306. https://doi.org/10.1073/pnas.1705895114
Bogaert, A. F., Skorska, M. N., Wang, C., Gabrie, J., MacNeil, A. J., Hoffarth, M. R., VanderLaan, D. P., Zucker, K. J., & Blanchard, R. (2022). Correction to supporting information for Bogaert et al., Male homosexuality and maternal immune responsivity to the Y-linked protein NLGN4Y. Proceedings of the National Academy of Sciences of the United States of America, 119, Article e2212767119. https://doi.org/10.1073/pnas.2212767119
Boyle, E. A., Li, Y. I., & Pritchard, J. K. (2017). An expanded view of complex traits: From polygenic to omnigenic. Cell, 169, 1177–1186. https://doi.org/10.1016/j.cell.2017.05.038
Byne, W., Tobet, S., Mattiace, L. A., Lasco, M. S., Kemether, E., Edgar, M. A., Morgello, S., Buchsbaum, M. S., & Jones, L. B. (2001). The interstitial nuclei of the human anterior hypothalamus: An investigation of variation with sex, sexual orientation, and HIV status. Hormones and Behavior, 40, 86–92. https://doi.org/10.1006/hbeh.2001.1680
Camperio Ciani, A., & Pellizzari, E. (2012). Fecundity of paternal and maternal non-parental female relatives of homosexual and heterosexual men. PLoS ONE, 7, Article e51088. https://doi.org/10.1371/journal.pone.0051088
Camperio Ciani, A., Cermelli, P., & Zanzotto, G. (2008). Sexually antagonistic selection in human male homosexuality. PLoS ONE, 3, Article e2282. https://doi.org/10.1371/journal.pone.0002282
Camperio Ciani, A., Corna, F., & Capiluppi, C. (2004). Evidence for maternally inherited factors favouring male homosexuality and promoting female fecundity. Proceedings of the Royal Society B: Biological Sciences, 271, 2217–2221.
Cantor, J. M., Blanchard, R., Paterson, A. D., & Bogaert, A. F. (2002). How many gay men owe their sexual orientation to fraternal birth order? Archives of Sexual Behavior, 31, 63–71. https://doi.org/10.1023/A:1014031201935
Cinelli, C., & Hazlett, C. (2020). Making sense of sensitivity: Extending omitted variable bias. Journal of the Royal Statistical Society: Series B, 82, 39–67. https://doi.org/10.1111/rssb.12348
Clive, J., Flintham, E., & Savolainen, V. (2023). Same-sex sociosexual behaviour is widespread and heritable in male rhesus macaques. Nature Ecology & Evolution, 7, 1287–1301. https://doi.org/10.1038/s41559-023-02111-y
Felesina, T. (2026). Understanding the evolutionary history of human male and female same-sex sexual behavior requires a greater appreciation of its shared genetic architecture. Archives of Sexual Behavior, 55, 1451–1453. https://doi.org/10.1007/s10508-026-03478-2
Felesina, T., & Zietsch, B. P. (2024). The desirable dad hypothesis: Male same-sex attraction as the product of selection for paternal care via antagonistic pleiotropy. Archives of Sexual Behavior, 53, 1731–1745.
Finucane, H. K., Bulik-Sullivan, B., Gusev, A., Trynka, G., Reshef, Y., Loh, P.-R., Anttila, V., Xu, H., Zang, C., Farh, K., Ripke, S., Day, F. R., Purcell, S., Stahl, E., Lindstrom, S., Perry, J. R. B., Okada, Y., Raychaudhuri, S., Daly, M. J., … Price, A. L. (2015). Partitioning heritability by functional annotation using genome-wide association summary statistics. Nature Genetics, 47, 1228–1235. https://doi.org/10.1038/ng.3404
Fořt, J., Flegr, J., Kuba, R., & Kaňková, Š. (2024). Fertility of Czech gay and straight men, women, and their relatives: Testing the sexually antagonistic gene hypothesis. Archives of Sexual Behavior, 53, 1747–1761.
Frigerio, A., Ballerini, L., & Valdés Hernández, M. (2021). Structural, functional, and metabolic brain differences as a function of gender identity or sexual orientation: A systematic review of the human neuroimaging literature. Archives of Sexual Behavior, 50, 3329–3352. https://doi.org/10.1007/s10508-021-02005-9
Ganna, A., Verweij, K. J. H., Nivard, M. G., Maier, R., Wedow, R., Busch, A. S., Abdellaoui, A., Guo, S., Sathirapongsasuti, J. F., 23andMe Research Team, Lichtenstein, P., Lundström, S., Långström, N., Auton, A., Harris, K. M., Beecham, G. W., Martin, E. R., Sanders, A. R., Perry, J. R. B., … Zietsch, B. P. (2019). Large-scale GWAS reveals insights into the genetic architecture of same-sex sexual behavior. Science, 365, Article eaat7693. https://doi.org/10.1126/science.aat7693
Ganna, A., Verweij, K. J. H., Nivard, M. G., Maier, R., Wedow, R., Busch, A. S., Abdellaoui, A., Guo, S., Sathirapongsasuti, J. F., 23andMe Research Team, Lichtenstein, P., Lundström, S., Långström, N., Auton, A., Harris, K. M., Beecham, G. W., Martin, E. R., Sanders, A. R., Perry, J. R. B., … Zietsch, B. P. (2021). Response to comment on Large-scale GWAS reveals insights into the genetic architecture of same-sex sexual behavior. Science, 371, Article eaba5693. https://doi.org/10.1126/science.aba5693
Gavrilets, S., & Rice, W. R. (2006). Genetic models of homosexuality: Generating testable predictions. Proceedings of the Royal Society B: Biological Sciences, 273, 3031–3038. https://doi.org/10.1098/rspb.2006.3684
Gómez, J. M., González-Megías, A., & Verdú, M. (2023). The evolution of same-sex sexual behaviour in mammals. Nature Communications, 14, Article 5719. https://doi.org/10.1038/s41467-023-41290-x
Hamer, D. H., Hu, S., Magnuson, V. L., Hu, N., & Pattatucci, A. M. L. (1993). A linkage between DNA markers on the X chromosome and male sexual orientation. Science, 261, 321–327.
Hamer, D. H., Mustanski, B., Sell, R., Sanders, S. A., & Garcia, J. R. (2021). Comment on Large-scale GWAS reveals insights into the genetic architecture of same-sex sexual behavior. Science, 371, Article eaba2941. https://doi.org/10.1126/science.aba2941
Hernán, M. A., & Robins, J. M. (2020). Causal inference: What if. Chapman & Hall/CRC.
Hines, M. (2011). Prenatal endocrine influences on sexual orientation and on sexually differentiated childhood behavior. Frontiers in Neuroendocrinology, 32, 170–182. https://doi.org/10.1016/j.yfrne.2011.02.006
Hu, S., Pattatucci, A. M. L., Patterson, C., Li, L., Fulker, D. W., Cherny, S. S., Kruglyak, L., & Hamer, D. H. (1995). Linkage between sexual orientation and chromosome Xq28 in males but not in females. Nature Genetics, 11, 248–256. https://doi.org/10.1038/ng1195-248
Hu, S.-H., Li, H.-M., Yu, H., Liu, Y., Liu, C.-X., Zuo, X.-B., Lu, J., Jiang, J.-J., Xi, C.-X., Huang, B.-C., Xu, H.-J., Hu, J.-B., Lai, J.-B., Huang, M.-L., Liu, J.-N., Xu, D.-G., Guo, X.-C., Wu, W., Wu, X., … Xu, Y. (2021). Discovery of new genetic loci for male sexual orientation in Han population. Cell Discovery, 7, Article 103. https://doi.org/10.1038/s41421-021-00341-7
Iemmola, F., & Camperio Ciani, A. (2009). New evidence of genetic factors influencing sexual orientation in men: Female fecundity increase in the maternal line. Archives of Sexual Behavior, 38, 393–399. https://doi.org/10.1007/s10508-008-9381-6
Imai, K., Keele, L., & Tingley, D. (2010). A general approach to causal mediation analysis. Psychological Methods, 15, 309–334. https://doi.org/10.1037/a0020761
Kendler, K. S., Thornton, L. M., Gilman, S. E., & Kessler, R. C. (2000). Sexual orientation in a U.S. national sample of twin and nontwin sibling pairs. American Journal of Psychiatry, 157, 1843–1846. https://doi.org/10.1176/appi.ajp.157.11.1843
Kim, K., Doi, A., Wen, B., Ng, K., Zhao, R., Cahan, P., Kim, J., Aryee, M. J., Ji, H., Ehrlich, L. I. R., Yabuuchi, A., Takeuchi, A., Cunniff, K. C., Hongguang, H., McKinney-Freeman, S., Naveiras, O., Yoon, T. J., Irizarry, R. A., Jung, N., … Daley, G. Q. (2010). Epigenetic memory in induced pluripotent stem cells. Nature, 467, 285–290. https://doi.org/10.1038/nature09342
Lakens, D., Scheel, A. M., & Isager, P. M. (2018). Equivalence testing for psychological research: A tutorial. Advances in Methods and Practices in Psychological Science, 1, 259–269. https://doi.org/10.1177/2515245918770963
Långström, N., Rahman, Q., Carlström, E., & Lichtenstein, P. (2010). Genetic and environmental effects on same-sex sexual behavior: A population study of twins in Sweden. Archives of Sexual Behavior, 39, 75–80. https://doi.org/10.1007/s10508-008-9386-1
Lerch, B. A. (2026). The evolution of same-sex sexual behavior: Using old theory to answer new questions. Integrative and Comparative Biology, 66, Article icag015. https://doi.org/10.1093/icb/icag015
LeVay, S. (1991). A difference in hypothalamic structure between heterosexual and homosexual men. Science, 253, 1034–1037.
Miller, E. M. (2000). Homosexuality, birth order, and evolution: Toward an equilibrium reproductive economics of homosexuality. Archives of Sexual Behavior, 29, 1–34.
Monk, J. D., Giglio, E., Kamath, A., Lambert, M. R., & McDonough, C. E. (2019). An alternative hypothesis for the evolution of same-sex sexual behaviour in animals. Nature Ecology & Evolution, 3, 1622–1631. https://doi.org/10.1038/s41559-019-1019-7
Mustanski, B. S., DuPree, M. G., Nievergelt, C. M., Bocklandt, S., Schork, N. J., & Hamer, D. H. (2005). A genomewide scan of male sexual orientation. Human Genetics, 116, 272–278. https://doi.org/10.1007/s00439-004-1241-4
National Academies of Sciences, Engineering, and Medicine. (2023). Using population descriptors in genetics and genomics research: A new framework for an evolving field. National Academies Press. https://doi.org/10.17226/26902
Ngun, T. C., Guo, W., Ghahramani, N. M., Purkayastha, K., Conn, D., Sanchez, F. J., Bocklandt, S., Zhang, M., Ramirez, C. M., Pellegrini, M., & Vilain, E. (2015, October 8). A novel predictive model of sexual orientation using epigenetic markers (Abstract No. 95) [Conference presentation]. American Society of Human Genetics 2015 Annual Meeting, Baltimore, MD, United States.
Pearl, J. (2009). Causality: Models, reasoning, and inference (2nd ed.). Cambridge University Press.
Phoenix, C. H., Goy, R. W., Gerall, A. A., & Young, W. C. (1959). Organizing action of prenatally administered testosterone propionate on the tissues mediating mating behavior in the female guinea pig. Endocrinology, 65, 369–382. https://doi.org/10.1210/endo-65-3-369
Rahman, Q., & Wilson, G. D. (2003). Born gay? The psychobiology of human sexual orientation. Personality and Individual Differences, 34, 1337–1382. https://doi.org/10.1016/S0191-8869(02)00140-X
Rice, G., Anderson, C., Risch, N., & Ebers, G. (1999). Male homosexuality: Absence of linkage to microsatellite markers at Xq28. Science, 284, 665–667. https://doi.org/10.1126/science.284.5414.665
Rice, W. R., Friberg, U., & Gavrilets, S. (2012). Homosexuality as a consequence of epigenetically canalized sexual development. The Quarterly Review of Biology, 87, 343–368.
Robins, J. M., & Greenland, S. (1992). Identifiability and exchangeability for direct and indirect effects. Epidemiology, 3, 143–155. https://doi.org/10.1097/00001648-199203000-00013
Roselli, C. E., Reddy, R. C., & Kaufman, K. R. (2011). The development of male-oriented behavior in rams. Frontiers in Neuroendocrinology, 32, 164–169. https://doi.org/10.1016/j.yfrne.2010.12.007
Sanders, A. R., Martin, E. R., Beecham, G. W., Guo, S., Dawood, K., Rieger, G., Badner, J. A., Gershon, E. S., Krishnappa, R. S., Kolundzija, A. B., Duan, J., Gejman, P. V., & Bailey, J. M. (2015). Genome-wide scan demonstrates significant linkage for male sexual orientation. Psychological Medicine, 45, 1379–1388. https://doi.org/10.1017/S0033291714002451
Semenyna, S. W., Gómez Jiménez, F. R., VanderLaan, D. P., & Vasey, P. L. (2023). Male androphilia, fraternal birth order, and female fecundity in Samoa: A 10-y retrospective. Proceedings of the National Academy of Sciences of the United States of America, 120, Article e2313284120. https://doi.org/10.1073/pnas.2313284120
Shapley, L. S. (1953). A value for n-person games. In H. W. Kuhn & A. W. Tucker (Eds.), Contributions to the theory of games (Vol. 2, pp. 307–317). Princeton University Press.
Song, S., & Zhang, J. (2023). Contraception ends the genetic maintenance of human same-sex sexual behavior. Proceedings of the National Academy of Sciences of the United States of America, 120, Article e2303418120. https://doi.org/10.1073/pnas.2303418120
Song, S., & Zhang, J. (2024). Genetic variants underlying human bisexual behavior are reproductively advantageous. Science Advances, 10, Article eadj6958. https://doi.org/10.1126/sciadv.adj6958
Swaab, D. F., & Hofman, M. A. (1990). An enlarged suprachiasmatic nucleus in homosexual men. Brain Research, 537, 141–148. https://doi.org/10.1016/0006-8993(90)90350-K
Swift-Gallant, A., Aung, T., Rosenfield, K., Dawood, K., & Puts, D. (2023). Organizational effects of gonadal hormones on human sexual orientation. Adaptive Human Behavior and Physiology, 9, 344–370. https://doi.org/10.1007/s40750-023-00226-x
VanderLaan, D. P., Skorska, M. N., Peragine, D. E., & Coome, L. A. (2022). Carving the biodevelopment of same-sex sexual orientation at its joints. In D. P. VanderLaan & W. I. Wong (Eds.), Gender and sexuality development: Contemporary theory and research (pp. 491–538). Springer. https://doi.org/10.1007/978-3-030-84273-4_17
VanderWeele, T. J. (2015). Explanation in causal inference: Methods for mediation and interaction. Oxford University Press.
VanderWeele, T. J., & Ding, P. (2017). Sensitivity analysis in observational research: Introducing the E-value. Annals of Internal Medicine, 167, 268–274. https://doi.org/10.7326/M16-2607
VanderWeele, T. J., Vansteelandt, S., & Robins, J. M. (2014). Effect decomposition in the presence of an exposure-induced mediator-outcome confounder. Epidemiology, 25, 300–306. https://doi.org/10.1097/EDE.0000000000000034
Vasey, P. L., & VanderLaan, D. P. (2010). Avuncular tendencies and the evolution of male androphilia in Samoan fa'afafine. Archives of Sexual Behavior, 39, 821–830. https://doi.org/10.1007/s10508-008-9404-3
Vilsmeier, J. K., Kossmeier, M., Voracek, M., & Tran, U. S. (2023). The fraternal birth-order effect as a statistical artefact: Convergent evidence from probability calculus, simulated data, and multiverse meta-analysis. PeerJ, 11, Article e15623.
Zietsch, B. P., Morley, K. I., Shekar, S. N., Verweij, K. J. H., Keller, M. C., Macgregor, S., Wright, M. J., Bailey, J. M., & Martin, N. G. (2008). Genetic factors predisposing to homosexuality may increase mating success in heterosexuals. Evolution and Human Behavior, 29, 424–433.
Zietsch, B. P., Sidari, M. J., Abdellaoui, A., Maier, R., Långström, N., Guo, S., Beecham, G. W., Martin, E. R., Sanders, A. R., & Verweij, K. J. H. (2021). Genomic evidence consistent with antagonistic pleiotropy may help explain the evolutionary maintenance of same-sex sexual behaviour in humans. Nature Human Behaviour, 5, 1251–1258.
Supplementary Material
The supplementary material (S1–S13, with Figures S1 and S2 and the full critical-study protocols) is included in the PDF after the references. Open the PDF.
Recommended Citation
Moore, T. M. (2026). The evolutionary persistence of male androphilia: A causal-portfolio theory of developmental paths and selection accounting. Manuscript retrieved from https://mooremetrics.com/homosexuality. https://doi.org/10.5281/zenodo.23789873